Class-wide sequence, structural, and docking-profile diversity of insect odorant receptors and structural specialization of Orco
Curation statements for this article:-
Curated by eLife

eLife Assessment
This important large-scale comparative study of odorant receptor (OR) genes across more than 100 insect species, combining sequence- and structure-based approaches, represents a significant contribution in the exploration of the evolution of this large gene family involved in the detection of odorant signals by olfactory neurons. Most notably, the authors uncover a structural feature unique to the odorant receptor co-receptor Orco that reduces ligand binding affinity. From this solid data emerges a general integrative framework to apprehend the diversity and evolution of odorant receptor genes in insects.
This article has been Reviewed by the following groups
Listed in
- Evaluated articles (eLife)
Abstract
Insect olfaction is facilitated by a heterotetrameric odorant receptor–odorant receptor co-receptor (OR-Orco) complex. However, extensive sequence divergence and lineage-specific expansion have complicated large-scale comparisons of OR repertoires across insects. Here, we integrated sequence similarity, structural similarity, and molecular docking to analyze ORs from 115 insect species. OR repertoire size was associated with adult diet, larval diet, and habitat, but not with circadian rhythm. We developed a ‘trunk–branch’ network framework to classify ORs according to sequence and structural similarity and used large-scale molecular docking to derive comparative docking profiles. Despite marked differences in sequence- and structural-community composition, almost all sampled orders contained ORs assigned to all six docking-profile communities. Holometabolous insects showed higher OR sequence-community diversity than non-holometabolous insects, and docking-profile composition differed between extant lineages whose order-level origins predate or postdate the end-Permian mass extinction. Most notably, we identified a highly conserved extracellular β-sheet in Orco that restricts ligand access to the binding pocket and may contribute to its reduced capacity to bind natural odorants. The emergence of Orco represents a key evolutionary transition from homomeric receptors to heteromeric OR–Orco complexes, and our results suggest that this transition may have been accompanied by pronounced specialization of the extracellular domain and binding pocket. Together, these analyses provide a class-wide framework for comparing insect OR diversity and generate testable hypotheses for future experimental and evolutionary studies.
Article activity feed
-

-
-

eLife Assessment
This important large-scale comparative study of odorant receptor (OR) genes across more than 100 insect species, combining sequence- and structure-based approaches, represents a significant contribution in the exploration of the evolution of this large gene family involved in the detection of odorant signals by olfactory neurons. Most notably, the authors uncover a structural feature unique to the odorant receptor co-receptor Orco that reduces ligand binding affinity. From this solid data emerges a general integrative framework to apprehend the diversity and evolution of odorant receptor genes in insects.
-
Reviewer #1 (Public review):
[Editors' note: this version has been assessed by the Reviewing Editor without further input from the original reviewers. All the reviewers' comments have been adequately addressed and taken into account.]
Objectives of the study and impact of the work
The authors of this article primarily aim to reconstruct the evolutionary history of the insect odorant receptor (OR) family, which is responsible for the detection of odorant signals by olfactory neurons. Due to the lack of phylogenetic signal present in the sequences of this multigene family, which evolves very rapidly, phylogenetic analyses have so far never made it possible to precisely retrace how ORs diversified prior to the appearance of present-day insect orders, and what the drivers of this diversification were. For example, one may suspect that the …
Reviewer #1 (Public review):
[Editors' note: this version has been assessed by the Reviewing Editor without further input from the original reviewers. All the reviewers' comments have been adequately addressed and taken into account.]
Objectives of the study and impact of the work
The authors of this article primarily aim to reconstruct the evolutionary history of the insect odorant receptor (OR) family, which is responsible for the detection of odorant signals by olfactory neurons. Due to the lack of phylogenetic signal present in the sequences of this multigene family, which evolves very rapidly, phylogenetic analyses have so far never made it possible to precisely retrace how ORs diversified prior to the appearance of present-day insect orders, and what the drivers of this diversification were. For example, one may suspect that the adaptation of ORs to odors emitted by plants constituted a critical step in insect evolution during the "angiosperm terrestrial revolution," which occurred at the end of the Cretaceous, but nothing currently allows this to be asserted.
There are very nice examples, notably in drosophilids, derived from comparisons between closely related species and documenting mechanisms of OR adaptation to certain signals. However, what the authors attempt to do in this work is to produce a macroevolutionary analysis at the scale of insects as a whole, based almost exclusively on bioinformatic analyses. To do this, they annotated OR genes in about one hundred insect species and developed pipelines for analyzing sequence similarity, structural similarity and functional similarity, the latter being estimated through a molecular docking approach. An important element in the evolution of insect ORs is the appearance of a unique co-receptor, called Orco, which appears to be an OR that has lost the ability to bind odorants. In addition to the large-scale bioinformatic analysis, the authors also aim to explore more specifically the factors that favored the emergence of Orco and the selective advantage conferred by the existence of OR-Orco complexes.
Given the importance of odorant receptors in insect biology and in their adaptation to different environments and lifestyles, retracing their evolutionary history is indeed a major question in evolutionary biology. In principle, this type of work therefore has the potential to become a reference in the field and to provide a basis for significant scientific advances.
Major strengths and weaknesses
The sampling chosen for collecting OR sequences is very impressive, with more than 100 insect families represented, covering most of the major orders. This sampling appears appropriate for the question being addressed. The analysis pipeline used to collect the sequences makes sense, relying on homology-based annotation tools coupled with a structure-based filter. Nevertheless, one can note aberrant numbers of ORs for certain species (much lower than reality). A lower number of OR genes is somewhat expected, as the authors chose to apply a fairly stringent filter on sequence quality (based on predicted 3D structure), which reduces the number from 14,000 to 9,000. This choice seems logical given the subsequent use of these data, but it inevitably leads to data loss. However, the low number of genes also results from the fact that the pipeline did not function correctly for all genomes.
In the revised version of the manuscript, the authors included a benchmarking step, which is a good point. They compared their OR gene annotations with previous reports in the same species. Unfortunately, this comparison is irrelevant because the chosen reference OR repertoires are actually a mix of transcriptome and genome annotations. Furthermore, comparisons with entire OR gene repertoires essentially demonstrate that their annotation was of good quality for species in which OR sequences were already present in the query OR database, but poor for species in which they were not. Therefore, these supplementary analyses made by the authors are not particularly in favor of an overall good quality of OR gene repertoires in the >110 species studied. The fact that some OR genes may be missing and that the total number may not be exact for each species is not prohibitive for studying the evolution of the family on a broad scale. However, it does call into question the correlation between the number of ORs and lifestyles and diets.
From the dataset collected, the authors attempted to categorize ORs in several ways, starting with the reconstruction of sequence similarity networks. The approach is interesting, but fails to reveal homology relationships between ORs from species belonging to different insect orders. So, it is unclear what the advantage of this approach is compared with the "classical" phylogenetic approach.
The clustering based on structure also leads to the identification of a majority of "order-specific" clusters, which does not provide major insights into the evolution of ORs. However, the authors highlight a group of ORs in flies that appear to possess an unusual intracellular region, as well as a cluster of OR shared across many insect orders that exhibit a larger binding cavity. This is really interesting, although more relevant to OR structure than to their evolution.
The analysis of structural diversity then leads the authors to focus on the Orco co-receptors, which are characterized by modifications of the binding pocket and the appearance of an extracellular loop that could explain the loss of the ability to bind odorant molecules. This part, which relies on in vitro experiments, is interesting and constitutes the most striking result of this study, which could in itself have been the subject of a separate manuscript.
The rest of the manuscript is based on the prediction of OR response spectra using molecular docking. The work that has been carried out is extremely substantial, and the objective of linking clusters based on sequence similarity or 3D structural similarity with functional categories is entirely relevant. The docking score threshold used was chosen thoughtfully, which is very good, and according to the calculation performed should ensure a true positive rate of more than 20%, which is excellent in such a docking analysis. But in the absence of functional validation, this 20% true positive rate is not sufficient to extrapolate OR function, and docking-derived binding breadth measures used in the remaining of the manuscript have to be taken with caution, as acknowledged by the authors themselves. Consequently, the chance that results of this part of the work will enlighten the evolution of OR on a broad scale is low. For example, the fact that insect lineages that emerged after the Permian-Triassic extinction have more broadly-tuned OR is an interesting observation, yet remains highly speculative.
In summary, despite the large number of analyses performed, the authors do not really succeed in achieving the stated objective of reconstructing the evolutionary history of insect ORs, and the results obtained do not strongly support all the conclusions regarding the links between OR repertoires and environment or lifestyle.
-
Reviewer #2 (Public review):
The remarkable evolvability of the olfactory system enables animals to rapidly adapt to dynamic and chemically complex environments. Over the past two decades, substantial effort has been devoted to uncovering the evolutionary principles that drive the diversification of odorant receptors (ORs), yielding key insights into the forces shaping their striking variability in both vertebrates and insects. In this manuscript, Zhang and colleagues analyze the OR repertoires of over 100 insect species, leveraging sequence and structural similarity to infer patterns of gene family evolution within this diverse and ecologically important clade. By integrating sequence-based and structure-based comparisons, their study builds on a compelling and recently emerging line of research made possible by the advent of …
Reviewer #2 (Public review):
The remarkable evolvability of the olfactory system enables animals to rapidly adapt to dynamic and chemically complex environments. Over the past two decades, substantial effort has been devoted to uncovering the evolutionary principles that drive the diversification of odorant receptors (ORs), yielding key insights into the forces shaping their striking variability in both vertebrates and insects. In this manuscript, Zhang and colleagues analyze the OR repertoires of over 100 insect species, leveraging sequence and structural similarity to infer patterns of gene family evolution within this diverse and ecologically important clade. By integrating sequence-based and structure-based comparisons, their study builds on a compelling and recently emerging line of research made possible by the advent of AlphaFold, which has previously clarified the phylogenetic relationship between insect Ors and the gustatory receptor gene family and revealed the unexpectedly deep evolutionary origins of this ancient structural fold.
Applying this approach to a large set of ORs derived from species throughout the insect phylogeny, the authors confirm many previously reported patterns of OR evolution. More importantly, with their large-scale approach the authors found new structural features of different Or clades, resulting in intriguing testable hypotheses about Or evolution that have the potential to support future directions in studying this important gene family. For example, the authors identify a structural feature mostly unique to the OR co-receptor ORco, a beta-sheet in EL2, which they functionally show reduces odorant binding affinity - a key aspect of ORco, which does not bind ligands in the ancestral ligand-biding site. This is a particularly strong part of the manuscript, since the authors support their in silico-derived hypothesis with functional data.
In an attempt to assess the relationship between sequence identity and structure on one hand and function on the other, the authors perform an in-silico structure prediction and chemical docking analysis. While not suitable to assess the actual function of individual receptors without functional testing, this strategy revealed potential receptor clades with differences in the number and type of docked chemicals. As a response to the previous reviews, the authors now changed their writing to refer to these results as 'docking-derived function' instead of just 'function'. While I appreciate that the authors changed their wording, this semantic change still can be read as referring to docking-derived predictions as conclusions of actual biological function. However, this is not the case. Any functional claims based solely on the docking approach need to be interpreted with caution, as in silico-only approaches are currently not well enough predictors for actual receptor function. Accordingly, I strongly recommend to remove the parts that connect receptor function to structure or ecology.
Assessing their docking analyses with respect to species ecology, the authors identify correlations suggesting that Or repertoires might follow predictable evolutionary trajectories broadly aligned with species ecology. These are interesting hypotheses. However, due to the inability to predict actual receptor function the author's claims about ecology-function relationships are not well supported. Accordingly, the authors should change the respective parts of the manuscript in way that it is clear these are currently untested hypotheses. This includes adjusting the title, since the work does not find any experimental support for ecological tuning.
I believe that future studies combining functional testing with an explicit evo-eco framework could benefit from this work and the explicit hypotheses it generates. But the authors should be careful to not overstate their in-silico derived results.
-
Author response:
The following is the authors’ response to the previous reviews.
Public Reviews:
Reviewer #1 (Public review):
Objectives of the study and impact of the work
The authors of this article primarily aim to reconstruct the evolutionary history of the insect odorant receptor (OR) family, which is responsible for the detection of odorant signals by olfactory neurons. Due to the lack of phylogenetic signal present in the sequences of this multigene family, which evolves very rapidly, phylogenetic analyses have so far never made it possible to precisely retrace how ORs diversified prior to the appearance of present-day insect orders, and what the drivers of this diversification were. For example, one may suspect that the adaptation of ORs to odors emitted by plants constituted a critical step in insect evolution during the …
Author response:
The following is the authors’ response to the previous reviews.
Public Reviews:
Reviewer #1 (Public review):
Objectives of the study and impact of the work
The authors of this article primarily aim to reconstruct the evolutionary history of the insect odorant receptor (OR) family, which is responsible for the detection of odorant signals by olfactory neurons. Due to the lack of phylogenetic signal present in the sequences of this multigene family, which evolves very rapidly, phylogenetic analyses have so far never made it possible to precisely retrace how ORs diversified prior to the appearance of present-day insect orders, and what the drivers of this diversification were. For example, one may suspect that the adaptation of ORs to odors emitted by plants constituted a critical step in insect evolution during the "angiosperm terrestrial revolution," which occurred at the end of the Cretaceous, but nothing currently allows this to be asserted.
There are very nice examples, notably in drosophilids, derived from comparisons between closely related species and documenting mechanisms of OR adaptation to certain signals. However, what the authors attempt to do in this work is to produce a macroevolutionary analysis at the scale of insects as a whole, based almost exclusively on bioinformatic analyses. To do this, they annotated OR genes in about one hundred insect species and developed pipelines for analyzing sequence similarity, structural similarity and functional similarity, the latter being estimated through a molecular docking approach. An important element in the evolution of insect ORs is the appearance of a unique co-receptor, called Orco, which appears to be an OR that has lost the ability to bind odorants. In addition to the large-scale bioinformatic analysis, the authors also aim to explore more specifically the factors that favored the emergence of Orco and the selective advantage conferred by the existence of OR-Orco complexes.
Given the importance of odorant receptors in insect biology and in their adaptation to different environments and lifestyles, retracing their evolutionary history is indeed a major question in evolutionary biology. In principle, this type of work therefore has the potential to become a reference in the field and to provide a basis for significant scientific advances.
Major strengths and weaknesses
The sampling chosen for collecting OR sequences is very impressive, with more than 100 insect families represented, covering most of the major orders. This sampling appears appropriate for the question being addressed. The analysis pipeline used to collect the sequences makes sense, relying on homology-based annotation tools coupled with a structure-based filter. Nevertheless, one can note aberrant numbers of ORs for certain species (much lower than reality). A lower number of OR genes is somewhat expected, as the authors chose to apply a fairly stringent filter on sequence quality (based on predicted 3D structure), which reduces the number from 14,000 to 9,000. This choice seems logical given the subsequent use of these data, but it inevitably leads to data loss. However, the low number of genes also results from the fact that the pipeline did not function correctly for all genomes.
In the revised version of the manuscript, the authors included a benchmarking step, which is a good point. They compared their OR gene annotations with previous reports in the same species. Unfortunately, this comparison is irrelevant because the chosen reference OR repertoires are actually a mix of transcriptome and genome annotations. Furthermore, comparisons with entire OR gene repertoires essentially demonstrate that their annotation was of good quality for species in which OR sequences were already present in the query OR database, but poor for species in which they were not. Therefore, these supplementary analyses made by the authors are not particularly in favor of an overall good quality of OR gene repertoires in the >110 species studied. The fact that some OR genes may be missing and that the total number may not be exact for each species is not prohibitive for studying the evolution of the family on a broad scale. However, it does call into question the correlation between the number of ORs and lifestyles and diets.
Thank you for this important comment. We agree that it is inappropriate to directly compare genome-based OR annotations with OR repertoires derived solely from transcriptomes, because transcriptome-only datasets generally do not capture the complete OR gene complement of a species. We therefore repeated the benchmarking analysis and collected, wherever available for the species included in this study, published OR repertoires based on genome annotation or genome annotation supported by transcriptomic evidence. Transcriptome-only OR datasets are no longer included in this comparison.
The updated benchmarking analysis shows that, for most species with comparable genome-based reference repertoires, the number of ORs identified in this study exceeds 70% of the previously reported number, with good agreement in OR repertoire size for most species (Table S1). Diabrotica virgifera is an exception: a previous manual annotation reported 124 complete ORs, whereas we identified 78, approximately two-thirds of the previously reported number. This result indicates that our automated annotation and stringent structural filtering pipeline may indeed underestimate OR repertoire size in some species. The large, repeat-rich genome of this species may further complicate gene prediction and the reconstruction of OR loci. We have therefore explicitly stated in the revised manuscript that incorporating transcriptomic evidence and manual curation may help recover more complete OR repertoires from such complex genomes.
Another notable outlier is Anagrus nilaparvatae. A previous study reported 88 ORs, whereas we identified 330. Examination of the methods used in that study revealed that its OR search relied primarily on homology searches using known OR sequences from two other hymenopteran species. Because insect ORs constitute a rapidly evolving gene family with pronounced lineage-specific expansions, a limited query set may fail to detect highly divergent or species-specific ORs. In comparison, our study used OR queries from a broader range of insects and a genome-level annotation pipeline specifically designed for insect ORs, which may have enabled the detection of more highly divergent candidate ORs. Because the protein sequences of the 88 ORs were not made available in the original study, we are currently unable to perform a one-to-one comparison to verify whether all of these ORs are included in our annotations.
You also noted that our better performance in species such as Drosophila melanogaster, Anopheles gambiae, and Bombyx mori might partly reflect the inclusion of known OR sequences from these species in the query database. We consider this an important and directly testable question. We therefore divided the benchmark species into two groups according to whether conspecific OR sequences were represented in the query database, and used ln(OR count_this study / OR count_previous) to quantify differences in OR counts relative to previous genome-based annotations. No significant difference was detected between the two groups (P = 0.841, Fig. S1 B). Given the unusually large difference in OR counts between the two studies for A. nilaparvatae, we also performed a sensitivity analysis excluding this species, which yielded the same conclusion (P = 0.413, Fig.S1 C). Thus, among the species for which comparable genome-based reference repertoires are currently available, we found no evidence that the inclusion of conspecific ORs in the query database systematically affected the final OR repertoire size. We have added this analysis to the supplementary material of the revised manuscript.
Finally, we agree that annotation biases in OR repertoire size may affect analyses of its correlations with ecological traits. We therefore repeated all relevant ecological sensitivity analyses using an updated OR count dataset, taking the larger of the OR counts from our study and the previous annotation for each species (Fig. S1 D- G). The main statistical results remained consistent with the original analyses: OR counts were still significantly associated with adult diet, larval diet, and habitat, whereas no significant association was detected with circadian rhythm. We therefore consider the statistical associations between OR counts and the main ecological variables to be reasonably robust within the range of annotation discrepancies detectable using the currently available genome-based benchmarks, although automated annotation and stringent structural filtering may underestimate OR counts in some species. The updated benchmarking analyses are described in Supplementary Text S1 and summarized in Table S1.
The following revision has been added (lines 103-107): “Benchmarking against curated OR repertoires showed that the automated pipeline recovered most previously reported ORs and that stringent structural filtering produced a high-confidence set of structurally intact receptors (Supplementary Text S1, Tables S1–S3, and fig. S1B–C). For most species with comparable genome-based references, retained OR counts exceeded 70% of published values.”
From the dataset collected, the authors attempted to categorize ORs in several ways, starting with the reconstruction of sequence similarity networks. The approach is interesting, but fails to reveal homology relationships between ORs from species belonging to different insect orders. So it is unclear what the advantage of this approach is compared with the "classical" phylogenetic approach.
Thank you for raising this important question. Our intention was not to replace classical phylogenetic analysis with sequence similarity networks. Rather, we used network-based approaches as a class-wide framework to classify ORs independently according to sequence similarity, structural similarity, and docking profiles, and then compared how these different classifications correspond across insect taxa and major macroevolutionary transitions.
For smaller gene families with limited sequence divergence, defining subfamilies based on gene trees is a well-established and highly effective approach, because phylogenetic trees can both identify subfamilies and reveal their evolutionary relationships. However, the insect OR superfamily exhibits pronounced birth-and-death evolution, with frequent gene duplications and losses across lineages, and comprises a vast number of highly divergent sequences. Consequently, when a single phylogenetic tree is constructed for the entire OR superfamily, deep nodes often lack sufficient support, and relationships among subfamilies may be difficult to resolve consistently. It is therefore challenging to obtain a stable and consistent classification of thousands of ORs into subfamilies across insects using a single gene tree alone. We therefore used sequence similarity networks as a complementary classification approach. Their main advantage is that they allow large datasets to be grouped according to sequence similarity among all ORs without requiring the resolution of every deep phylogenetic relationship. More importantly, similar network frameworks can be used to construct sequence similarity, structural similarity, and docking-profile similarity networks, allowing SeqC, StrC, and dPC to be compared directly within a common analytical framework. This is difficult to achieve using conventional gene trees alone and is the main reason for our choice of a network-based approach.
Sequence similarity networks alone cannot directly resolve homology relationships among ORs from different insect orders. During the previous revision, we therefore sought to link sequence communities to phylogenetic relationships. We conducted additional phylogenetic analyses of shared sequence communities containing multiple insect orders to search for potentially conserved OR lineages spanning different orders. However, among these shared communities, we identified only one conserved OR lineage spanning four insect orders with relatively clear phylogenetic support. We have added this result to the Supplementary Data S6. This finding further indicates that clearly identifiable orthology relationships spanning multiple orders are very limited among extant insect ORs.
The following revision has also been added (lines 150-154): “The SSN is not intended to replace phylogenetic reconstruction, but rather to provide a complementary framework for summarizing sequence relationships among thousands of highly divergent ORs at a class-wide scale. It allows major patterns of OR sequence similarity and diversification to be visualized more clearly across the entire dataset. Phylogenetic analyses can then be used to further resolve evolutionary relationships within specific sequence communities.”
The clustering based on structure also leads to the identification of a majority of "orderspecific" clusters, which does not provide major insights into the evolution of ORs. However, the authors highlight a group of ORs in flies that appear to possess an unusual intracellular region, as well as a cluster of OR shared across many insect orders that exhibit a larger binding cavity. This is really interesting, although more relevant to OR structure than to their evolution.
Thank you for this important comment. We agree that the previous version of the manuscript focused primarily on structural differences among structural communities and did not adequately explore their potential evolutionary implications. StrC23 is the most widely distributed structural community, spanning 12 insect orders and containing 46.3% of the ORs. We further found that its putative binding pockets were significantly larger than those of other ORs, which we previously interpreted as a potentially conserved structural pattern across taxa. Following your suggestion, we conducted additional analyses of the phylogenetic and orthology relationships within StrC23. The results showed that StrC23 does not correspond to a single, broadly conserved orthogroup spanning insect orders, but instead contains 156 distinct orthologous groups. Of these, 24 encompass more than 10 species. Only a few of these larger groups span multiple insect orders: one contains ORs from Hymenoptera, Diptera, and Trichoptera, and two contain ORs from both Lepidoptera and Trichoptera. The remaining groups show pronounced order specificity, including nine Hymenoptera-specific groups, eight Lepidoptera-specific groups, and four Diptera-specific groups.
These results indicate that the large putative binding pocket shared by StrC23 members cannot simply be attributed to a single conserved orthologous lineage spanning multiple orders, but is distributed across several phylogenetically independent OR lineages. This pattern may reflect convergent evolution toward similar structural features in different OR lineages, or the retention of an older structural feature in multiple OR lineages that subsequently underwent substantial sequence divergence. The current data do not allow us to distinguish conclusively between these two possibilities. We have added the phylogenetic and orthologous-group analyses of StrC23 to the revised manuscript and supplementary material to further clarify the evolutionary implications of this structural community shared across insect orders.
The following revision has been added (lines 208-218): “To further investigate the evolutionary basis of this broadly shared structural feature, we examined the phylogenetic and orthology relationships among StrC23 receptors. StrC23 was divided into 156 orthologous groups, 24 of which contained ORs from more than 10 species. However, only a few of these larger groups spanned multiple insect orders: one included ORs from Hymenoptera, Diptera, and Trichoptera, while two included ORs from both Lepidoptera and Trichoptera (Supplementary Data S6). Thus, StrC23 does not represent a single conserved cross-order orthologous lineage, but instead comprises multiple phylogenetically distinct OR lineages. Its shared large-pocket architecture therefore cannot be explained solely by conservation of a single orthologous lineage and may reflect either structural convergence or retention of an ancient structural feature followed by substantial sequence divergence.”
The analysis of structural diversity then leads the authors to focus on the Orco co-receptors, which are characterized by modifications of the binding pocket and the appearance of an extracellular loop that could explain the loss of the ability to bind odorant molecules. This part, which relies on in vitro experiments, is interesting and constitutes the most striking result of this study, which could in itself have been the subject of a separate manuscript.
We thank the reviewer for the positive assessment of this part of our work.
The rest of the manuscript is based on the prediction of OR response spectra using molecular docking. The work that has been carried out is extremely substantial, and the objective of linking clusters based on sequence similarity or 3D structural similarity with functional categories is entirely relevant. The docking score threshold used was chosen thoughtfully, which is very good, and according to the calculation performed should ensure a true positive rate of more than 20%, which is excellent in such a docking analysis. But in the absence of functional validation, this 20% true positive rate is not sufficient to extrapolate OR function, and docking-derived binding breadth measures used in the remaining of the manuscript have to be taken with caution, as acknowledged by the authors themselves. Consequently, the chance that results of this part of the work will enlighten the evolution of OR on a broad scale is low. For example, the fact that insect lineages that emerged after the Permian-Triassic extinction have more broadly-tuned OR is an interesting observation, yet remains highly speculative.
Thank you for this important comment. We agree that, even after optimizing the docking threshold, the current hit rate of approximately 20% remains insufficient to interpret molecular docking results as reliable predictions of actual OR ligand responses or functions. We also acknowledge that the biological implications of some docking results were still overstated in the previous revision, potentially leading readers to interpret these computational results as approximations of actual OR function. To avoid this misunderstanding, we have further revised the relevant terminology and interpretation of results throughout the latest manuscript. For results derived entirely from molecular docking, we no longer use terms such as ‘docking-derived function’, ‘functional community’, or other expressions that might imply experimental evidence of function. Instead, we consistently use ‘docking profile’ to make clear that these analyses reflect only similarities in computational docking patterns among receptors.
In addition, following your comments and the related suggestions from Reviewer 2, we have removed all results and discussion that directly linked docking-based predictions of OR function or recognition breadth to species ecology. We have explicitly presented the few remaining macroevolutionary comparisons as hypothesis-generating observations rather than evidence of functional evolution. For example, differences in docking-profile composition among extant insect lineages that originated before or after the end-Permian mass extinction (EPME) are now described only as a broad statistical pattern, with any interpretation framed as an experimentally untested hypothesis. Whether this pattern reflects actual differences in receptor tuning, and whether it is evolutionarily linked to environmental changes associated with the EPME, still requires testing through broader taxonomic sampling, reliable ancestral-state reconstruction, and functional experiments. We hope that these revisions restrict the interpretation of molecular docking results to what the evidence can support and bring the corresponding conclusions into closer alignment with the current strength of evidence.
The following revision has been added in Result section (lines 367-369): “These observations should be interpreted as a docking-derived comparative pattern among extant lineages rather than direct evidence of ancestral functional changes across the
EPME.”
And Discussion section (lines 410-414): “However, this interpretation remains hypothetical. Because rapid OR evolution and limited cross-order orthology preclude reliable reconstruction of deep ancestral repertoires, the EPME-related pattern should be viewed as a descriptive comparison among extant lineages. Further validation will require broader taxon sampling, ancestral reconstruction, and experimental testing of receptor tuning breadth.”
In summary, despite the large number of analyses performed, the authors do not really succeed in achieving the stated objective of reconstructing the evolutionary history of insect ORs, and the results obtained do not strongly support all the conclusions regarding the links between OR repertoires and environment or lifestyle.
Thank you for this important comment. We agree that ‘reconstructing the evolutionary history of insect ORs’ was too broad a formulation and exceeded what our methods could reliably address. A rigorous reconstruction of evolutionary history requires well-resolved phylogenetic relationships across taxa, ancestral-state inference, and systematic analysis of gene duplication and loss events. We have therefore redefined the scientific objectives of this study. The revised manuscript no longer claims to have reconstructed the evolutionary history of insect ORs. Instead, it aims to systematically characterize insect OR diversity and classification patterns at the levels of sequence, structure, and docking profile, compare the correspondence among these levels, and describe differences in their distribution across insect taxa and broad macroevolutionary contexts.
We have also reassessed the strength of different types of evidence concerning the relationships between OR repertoires and environment or lifestyle. For relationships between OR repertoire size and ecological traits, we repeated the genome-based benchmarking. After rerunning the ecological correlation sensitivity analyses using the updated OR count dataset, the main statistical results remained consistent with the original analyses. We therefore consider the current data to continue to support statistical associations between OR counts and ecological traits, with a degree of robustness. By contrast, we have removed results and discussion directly linking predicted OR function or recognition breadth to species ecology, and no longer use terms such as ‘docking-derived function’ or ‘functional community’ that might imply experimental evidence of function. The remaining macroevolutionary comparisons of docking profiles, such as differences in docking-profile composition among groups considered in relation to the EPME, are treated only as hypothesis-generating comparative patterns, rather than direct evidence of receptor functional evolution. We explicitly state that these hypotheses require future testing through functional experiments and more reliable ancestral-state reconstruction.
In summary, we have redefined the study objectives and no longer describe this work as a reconstruction of insect OR evolutionary history. We have retained the statistical associations between OR counts and ecological traits that remained supported after reassessment, while removing results and discussion linking receptor function to ecology on the basis of molecular docking. We believe these revisions better align the scientific objectives and scope of the conclusions with the evidence provided by the available data and methods.
We have revised the final paragraph of the Introduction to clarify the scope and objectives of this study (lines 80-91): “In this study, we characterized large-scale patterns of OR diversification across 115 species spanning 16 insect orders. Using a framework that integrates network analysis, protein structure prediction, and molecular docking, we constructed sequence-similarity (SSN), structural-similarity (StSN), and docking-profile similarity networks (dPSN) to classify ORs from three distinct perspectives. We identified differences in OR sequence-community diversity between holometabolous and non-holometabolous insects, as well as differences in docking-profile community composition among extant lineages whose order-level origins fall on opposite sides of the end-Permian mass extinction (EPME). Across this class-wide sampling, OR repertoire size was associated with adult diet, larval diet, and habitat, but not with circadian rhythm. Moreover, the conserved β-sheet in the extracellular loop 2 (EL2) region of Orco, together with its specialized binding-pocket architecture, is associated with its reduced ligand-binding capacity and represents a prominent structural specialization of the co-receptor lineage.”
Reviewer #2 (Public review):
The remarkable evolvability of the olfactory system enables animals to rapidly adapt to dynamic and chemically complex environments. Over the past two decades, substantial effort has been devoted to uncovering the evolutionary principles that drive the diversification of odorant receptors (ORs), yielding key insights into the forces shaping their striking variability in both vertebrates and insects. In this manuscript, Zhang and colleagues analyze the OR repertoires of over 100 insect species, leveraging sequence and structural similarity to infer patterns of gene family evolution within this diverse and ecologically important clade. By integrating sequence-based and structure-based comparisons, their study builds on a compelling and recently emerging line of research made possible by the advent of AlphaFold, which has previously clarified the phylogenetic relationship between insect Ors and the gustatory receptor gene family and revealed the unexpectedly deep evolutionary origins of this ancient structural fold.
Applying this approach to a large set of ORs derived from species throughout the insect phylogeny, the authors confirm many previously reported patterns of OR evolution. More importantly, with their large-scale approach the authors found new structural features of different Or clades, resulting in intriguing testable hypotheses about Or evolution that have the potential to support future directions in studying this important gene family. For example, the authors identify a structural feature mostly unique to the OR co-receptor ORco, a beta-sheet in EL2, which they functionally show reduces odorant binding affinity - a key aspect of ORco, which does not bind ligands in the ancestral ligand-biding site. This is a particularly strong part of the manuscript, since the authors support their in silico-derived hypothesis with functional data.
We thank the reviewer for the positive assessment of this part of our work.
In an attempt to assess the relationship between sequence identity and structure on one hand and function on the other, the authors perform an in-silico structure prediction and chemical docking analysis. While not suitable to assess the actual function of individual receptors without functional testing, this strategy revealed potential receptor clades with differences in the number and type of docked chemicals. As a response to the previous reviews, the authors now changed their writing to refer to these results as 'docking-derived function' instead of just 'function'. While I appreciate that the authors changed their wording, this semantic change still can be read as referring to dockingderived predictions as conclusions of actual biological function. However, this is not the case. Any functional claims based solely on the docking approach need to be interpreted with caution, as in silico-only approaches are currently not well enough predictors for actual receptor function. Accordingly, I strongly recommend to remove the parts that connect receptor function to structure or ecology.
Thank you for highlighting this important issue again. We agree with your comments and acknowledge that the presentation of molecular docking results in the previous revision still overstated their biological implications. Following your suggestion, we have removed all results and discussion that directly linked receptor function to species ecology solely on the basis of molecular docking, to avoid interpreting computational predictions as actual biological functions. We then systematically reviewed and revised docking-related terminology throughout the manuscript. For results derived entirely from docking analyses, we no longer use ‘docking-derived function’, ‘functional community’, or other expressions that might imply experimental evidence of function. Instead, we consistently use ‘docking profile’ to clarify that these analyses reflect similarities in computational docking patterns among receptors, rather than experimentally validated functional categories.
Through these revisions, we hope to distinguish more clearly between computational docking profiles and experimentally validated receptor function, and to avoid biological interpretations of docking results that exceed the available evidence. We believe these changes bring the manuscript's conclusions into closer alignment with the strength of the evidence.
Assessing their docking analyses with respect to species ecology, the authors identify correlations suggesting that Or repertoires might follow predictable evolutionary trajectories broadly aligned with species ecology. These are interesting hypotheses. However, due to the inability to predict actual receptor function the author's claims about ecology-function relationships are not well supported. Accordingly, the authors should change the respective parts of the manuscript in way that it is clear these are currently untested hypotheses. This includes adjusting the title, since the work does not find any experimental support for ecological tuning.
Thank you for this important suggestion. We agree that the current molecular docking analyses cannot support direct conclusions about relationships between actual receptor function and species ecology. Following your suggestion, we have removed the results and discussion directly linking docking-based predictions of OR function to species ecology. For the remaining macroevolutionary comparisons, such as differences in docking-profile composition among groups considered in relation to the EPME, we explicitly state that these results represent only computational patterns observed in extant taxa. The evolutionary interpretations proposed from these patterns remain experimentally untested hypotheses and cannot be considered direct evidence of changes in actual receptor function.
In addition, following your suggestion, we have changed the manuscript title to ‘Classwide sequence, structural, and docking-profile diversity of insect odorant receptors and structural specialization of Orco’. The new title no longer emphasizes ‘ecological tuning’ or ‘olfactory perception’, but instead focuses on class-wide diversification of insect OR sequences, structures, and docking profiles, together with the experimentally supported structural specialization of Orco.
The following revision has been added in Result section (lines 367-369): “These observations should be interpreted as a docking-derived comparative pattern among extant lineages rather than direct evidence of ancestral functional changes across the EPME.”
And Discussion section (lines 410-414): “However, this interpretation remains hypothetical. Because rapid OR evolution and limited cross-order orthology preclude reliable reconstruction of deep ancestral repertoires, the EPME-related pattern should be viewed as a descriptive comparison among extant lineages. Further validation will require broader taxon sampling, ancestral reconstruction, and experimental testing of receptor tuning breadth.”
I believe that future studies combining functional testing with an explicit evo-eco framework could benefit from this work and the explicit hypotheses it generates. But the authors should be careful to not overstate their in-silico derived results.
Thank you for this suggestion. We have revised the manuscript accordingly.
Recommendations for the authors:
Reviewer #1 (Recommendations for the authors):
Below are my comments in response to the rebuttal letter. I thank the authors for their careful revision and detailed responses to the points raised in my review.
(1) OR annotation pipeline and correlation analyses
Previous comment: "One can note aberrant numbers of ORs for certain species (much lower than reality), which indicates that the pipeline probably did not function correctly for all genomes. In the absence of a validation step comparing the results with already known OR repertoires, it is difficult to estimate the overall quality of the data. [...] It calls into question certain results that rely on this total number, such as the correlation between the number of ORs and genome size, lifestyle, and diet."
The response made by the authors on that point is not particularly convincing. They compared their OR gene annotations with previous reports in the same species, but this comparison is irrelevant because the chosen reference OR repertoires are actually a mix of transcriptome and genome annotations (the OR counts for Harmonia axyridis, Plutella xylostella, Pierus brassicae, Dendrolimus kikuchii, Carposina sasakii, Hermetia illucens, Chlorops oryzae and Propsilocerus akamusi are from transcriptomes). As not all OR genes are expressed in a given tissue, the number of ORs in transcriptomes is always lower than in genomes. Furthermore, comparisons with entire OR gene repertoires essentially demonstrate that their annotation was of good quality for species in which OR sequences were already present in the query OR database (Bombyx mori, Drosophila melanogaster, Anopheles gambiae) but was of bad quality for species in which OR sequences were NOT in the query (Diabrotica virgifera). However, re-annotation of Diabrotica led to an OR repertoire of 78 complete genes, vs. 124 in the manually curated genome annotation (1/3rd of the genes are missing). So these supplementary analyses made by the authors are not particularly in favor of an overall good quality of OR gene repertoires in the >110 species studied. They have also repeated the correlation analyses using 'full' OR gene sets where available. This showed that the correlation of OR count with genome size is no longer valid, but the correlation with diet or lifestyle is. However, if a mix of gene numbers from transcriptomes and genomes was used (see my comment above), I am afraid that these novel correlation analyses are not relevant. In conclusion, I would disagree with the claim that "These new benchmark and sensitivity analyses directly strengthen the evidence supporting our dataset and its ecological interpretations".
Thank you for further highlighting this issue. As detailed in our response to the Public Review, we repeated the OR annotation benchmarking, excluded transcriptome-only repertoires, and restricted comparisons to genome-based reference datasets or genome-based datasets supported by transcriptomic evidence. The updated results show that, for most species with comparable reference data, the number of ORs identified in this study exceeds 70% of the previously reported number. We also explicitly acknowledge that repertoire size may still be underestimated in some species.
We also directly tested whether query database composition systematically affected the number of annotated ORs. No significant difference was detected between species with and without conspecific OR queries (P = 0.841; P = 0.413 after excluding Anagrus nilaparvatae). Thus, we currently find no evidence that conspecific queries systematically increase the final OR count.
In addition, we repeated the sensitivity analyses of ecological correlations using the updated OR count dataset. The main statistical results remained unchanged: OR counts were still significantly associated with adult diet, larval diet, and habitat, but not with circadian rhythm. We therefore consider these statistical associations between OR counts and ecological traits to be reasonably robust to the annotation differences currently detectable. Detailed results are provided in our response to the Public Review and in the revised supplementary text S1.
(2) sequence similarity-based clusters
Previous comment: "The authors attempted to categorize ORs in several ways, starting with the reconstruction of sequence similarity networks. The approach is interesting, but in the end, the results do not seem to be sufficiently exploited. [...] For example, clusters SeqC26 or SeqC28, which appear to be shared by many insects, are potentially very interesting. It might have been relevant to combine this similarity-based clustering approach with phylogenetic reconstructions within each shared cluster."
The authors have followed this suggestion and identified an orthogroup conserved across several insect orders, which is an interesting observation. However the phylogeny of this orthogroup should be provided in supplementary data, which does not seem to be the case.
We thank the reviewer for the positive assessment. The corresponding phylogenetic tree has been added to the supplementary data S6.
(3) structure-based clusters
Previous comment: "The clusters shared by several orders are not truly exploited, which does not provide new insight into the evolution of ORs"
The authors show an interesting feature for a cluster of ORs from several insect orders, with a large putative binding pocket. However, it is not clear whether they verified or not if this cluster also correspond to an orthogroup or if this shared feature (a large binding pocket) is a convergence. They also improved the discussion of their results on a cluster of fly ORs with a long intracellular domain, which is a good thing.
Thank you for this important question. To determine whether StrC23, a structural community shared across multiple insect orders, corresponds to a conserved orthogroup, we performed additional phylogenetic analyses of its OR sequences and identified orthology relationships within the community based on the gene tree. The results showed that StrC23 does not correspond to a single orthogroup spanning insect orders, but instead contains 156 distinct orthologous groups. Of these, 24 encompass more than 10 species. Among these 24 larger groups, one contains ORs from Hymenoptera, Diptera, and Trichoptera, and two contain ORs from both Lepidoptera and Trichoptera. The remaining groups show pronounced order specificity, including nine Hymenoptera-specific groups, eight Lepidoptera-specific groups, and four Diptera-specific groups.
These results indicate that the large putative binding pocket characteristic of StrC23 is not explained by a single, broadly conserved orthogroup spanning multiple orders, but is distributed across several phylogenetically independent OR lineages. We therefore consider that this shared structural feature cannot simply be interpreted as direct retention within a conserved orthologous lineage. It may reflect convergent evolution toward similar structural features in different OR lineages, or the retention of an older structural feature in multiple OR lineages that subsequently underwent substantial sequence divergence. The current data do not allow us to distinguish conclusively between these two possibilities. We have added the phylogenetic analysis of StrC23 to the revised manuscript to clarify the evolutionary relationships within this structural community.
The following revision has been added (lines 208-218): “To further investigate the evolutionary basis of this broadly shared structural feature, we examined the phylogenetic and orthology relationships among StrC23 receptors. StrC23 was divided into 156 orthologous groups, 24 of which contained ORs from more than 10 species. However, only a few of these larger groups spanned multiple insect orders: one included ORs from Hymenoptera, Diptera, and Trichoptera, while two included ORs from both Lepidoptera and Trichoptera (Supplementary Data S6). Thus, StrC23 does not represent a single conserved cross-order orthologous lineage, but instead comprises multiple phylogenetically distinct OR lineages. Its shared large-pocket architecture therefore cannot be explained solely by conservation of a single orthologous lineage and may reflect either structural convergence or retention of an ancient structural feature followed by substantial sequence divergence.”
(4) molecular dynamics simulation
Previous comment: "The molecular dynamics modelling does not add anything in the way it is conducted (5 ns is too short)."
The authors argue that such a short simulation is sufficient for testing the interaction between a compound and the extracellular beta-sheet of the coreceptor Orco. I'm not a specialist enough to confirm whether this is actually true, but I trust the author's expertise in this area.
We thank the reviewer for the positive assessment.
(5 / 6) molecular docking and functional extrapolation
Previous comment: "in the absence of functional validation, this 20% true positive rate is not sufficient to extrapolate OR function"
The authors argue that molecular docking was not used to identify specific OR/ligand pairs, but rather to "estimate the potential binding tendencies of OR repertoires". I agree with this point; correlations were made using "binding breadth index values" that are calculated for entire OR repertoires. The text of the manuscript has been edited to refer to "docking-derived" functional clusters and binding breadth index, and to emphasise that the results should be interpreted with caution, which is a good point as it will prevent potential misinterpretation by the reader. However, this does not change the fact that interpreting the olfactory capacity of one species based on docking-derived binding breadth values of its OR repertoire is highly speculative (especially when OR repertoires are incomplete) and the chance that results of this part of the work will enlighten the evolution of OR on a broad scale is low.
Thank you for raising this concern. We agree that repertoire-level analyses do not resolve the inherent limitations of molecular docking in predicting actual receptor function. Although such analyses summarize overall docking patterns rather than individual OR–ligand interactions, they still cannot reliably define a species’ true olfactory capabilities. It therefore remains insufficient to describe a species' actual olfactory capabilities solely on the basis of docking-derived binding breadth. As detailed in our response to the Public Review, we have addressed this issue by following your suggestions and those of Reviewer 2 and removing the results and discussion that used the binding breadth index to directly link predicted receptor function to ecological traits such as diet, habitat, and circadian rhythm. We have also further standardized the terminology, replacing ‘docking-derived function’, ‘functional community’, and other expressions that could imply experimentally established function with ‘docking profile’, to emphasize that these results describe computational docking patterns rather than experimentally determined receptor functions. Through these revisions, we hope to distinguish more clearly between computational docking patterns and experimentally established receptor functions, and to restrict the interpretation of the related macroevolutionary comparisons to what the current data can support.
(7) EPME
Previous comment: "ORs of non-holometabolous insects cannot be considered equivalent to those of species that existed before the Permian-Triassic extinction"
The authors have now clarified the fact that they compared extant insect orders rather than ancestral and derived OR repertoires.
We thank the reviewer for the positive assessment of the revised manuscript.
(8 to 16)
The authors have carefully considered all the points raised in the recommendations for the authors, improving the manuscript on several aspects.
We thank the reviewer for the positive assessment of the revised manuscript.
Minor comment
Line 217: Orco did not emerge in the ancestor of Zygentoma, it emerged in the common ancestor of Zygentoma and Pterygota.
Thank you for pointing this out. We have corrected the statement.
Reviewer #2 (Recommendations for the authors):
The authors have done a good job in their revision of the manuscript. They addressed many of the reviewers' concerns well.
We thank the reviewer for the positive assessment of our previous revision.
However, the functional claims are still too strong and need to be removed or tempered down as described in the public review.
Thank you for this suggestion. We have removed or tempered the functional predictions in the manuscript in accordance with the Public Review. Please see our response to the Public Review for details.
Further, the manuscript title is not a good fit for the content. I would argue that the paper does not 'reveal ecological tuning shaping olfactory perception', since Or structurefunction relationships are involved in olfactory sensation or detection. Perception, however, requires interpretation, which is done by neural circuits but not receptors expressed within. Further, I still think that referring to 'ecological tuning' is far-fetched based on the docking approach that cannot really resolve the tuning of receptor sets. I would think that the work reveals interesting correlations between docking-derived population tuning and ecology. But that does not necessarily have anything to do with actual function and thus may not make much sense to advertise in the title or abstract.
Thank you for your suggestions concerning the manuscript title. Following this comment, we have changed the title from ‘Evolutionary dynamics of insect odorant receptors reveal ecological tuning shaping olfactory perception’ to ‘Class-wide sequence, structural, and docking-profile diversity of insect odorant receptors and structural specialization of Orco’ The new title no longer emphasizes ecological tuning or olfactory perception, but instead focuses on the two aspects of this study supported by the most direct evidence: class-wide patterns of sequence and structural diversification in insect ORs, and the experimentally supported structural specialization of Orco.
Consistent with the revised title, we have further adjusted the relevant wording in the Abstract, Results, and Discussion, and removed content directly linking receptor function to species ecology on the basis of docking results. We believe that the revised title and abstract more accurately reflect the main content of the study and the scope of its evidence, while avoiding functional or ecological interpretations of computational predictions that exceed their evidential support.
I suggest selecting a title that fits the sequence/structure/in silico docking nature of their large macroevolutionary approach and findings better. After all, apart from the orco work this manuscript is mostly a resource of in silico predictions.
Thank you for this suggestion. We have revised the manuscript title.
Author response table 1.
Numbers of OR genes in previously annotated species.
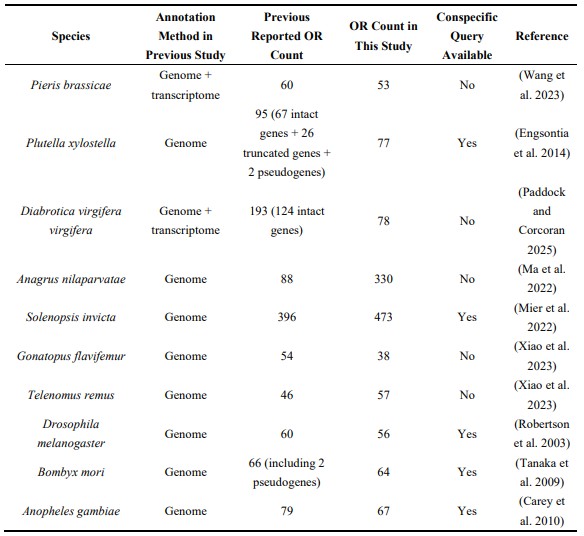
Reference
Carey AF, Wang G, Su CY, Zwiebel LJ, Carlson JR. 2010. Odorant reception in the malaria mosquito Anopheles gambiae. Nature 464: 66-71.
Engsontia P, Sangket U, Chotigeat W, Satasook C. 2014. Molecular evolution of the odorant and gustatory receptor genes in lepidopteran insects: implications for their adaptation and speciation. J Mol Evol 79: 21-39.
Ma Y, Guo Z, Wang L, Wang B, Huang T, Tang B, Zhang G, Zhou Q. 2022. The genome of the rice planthopper egg parasitoid wasp Anagrus nilaparvatae casts light on the chemo- and mechanosensation in parasitism. BMC Genomics 23: 541.
Mier P, Fontaine JF, Stoldt M, Libbrecht R, Martelli C, Foitzik S, Andrade-Navarro MA. 2022. Annotation and Analysis of 3902 Odorant Receptor Protein Sequences from 21 Insect Species Provide Insights into the Evolution of Odorant Receptor Gene Families in Solitary and Social Insects. Genes (Basel) 13.
Paddock KJ, Corcoran JA. 2025. Life-stage dependent behavior mimics chemosensory repertoire diversity in a belowground, specialist herbivore. G3 (Bethesda) 15.
Robertson HM, Warr CG, Carlson JR. 2003. Molecular evolution of the insect chemoreceptor gene superfamily in Drosophila melanogaster. Proc Natl Acad Sci U S A 100 Suppl 2: 14537-14542.
Tanaka K, Uda Y, Ono Y, Nakagawa T, Suwa M, Yamaoka R, Touhara K. 2009. Highly selective tuning of a silkworm olfactory receptor to a key mulberry leaf volatile. Curr Biol 19: 881-890.
Wang Q, Dicke M, Haverkamp A. 2023. Sympatric Pieris butterfly species exhibit a high conservation of chemoreceptors. Front Cell Neurosci 17: 1155405.
Xiao S, Ye X, Wang S, Yang Y, Fang Q, Wang F, Ye G. 2023. Genome assembly of the ectoparasitoid wasp Theocolax elegans. Sci Data 10: 159.
-
-

-
-
eLife Assessment
This valuable large-scale comparative study of odorant receptor (OR) genes across more than 100 insect species, combining sequence- and structure-based approaches, is a significant step in the exploration of the evolution of this large gene family involved in the detection of odorant signals by olfactory neurons. This work uncovers a structural feature unique to the odorant receptor co-receptor Orco that reduces ligand binding affinity. However, the strength of evidence is incomplete relative to the authors' claims about the inferred number of OR genes per species, the odor response spectra and the evolutionary and ecological interpretations.
-
Reviewer #1 (Public review):
Objectives of the study and impact of the work
The authors of this article primarily aim to reconstruct the evolutionary history of the insect odorant receptor (OR) family, which is responsible for the detection of odorant signals by olfactory neurons. Due to the lack of phylogenetic signal present in the sequences of this multigene family, which evolves very rapidly, phylogenetic analyses have so far never made it possible to precisely retrace how ORs diversified prior to the appearance of present-day insect orders, and what the drivers of this diversification were. For example, one may suspect that the adaptation of ORs to odors emitted by plants constituted a critical step in insect evolution during the "angiosperm terrestrial revolution," which occurred at the end of the Cretaceous, but nothing currently …
Reviewer #1 (Public review):
Objectives of the study and impact of the work
The authors of this article primarily aim to reconstruct the evolutionary history of the insect odorant receptor (OR) family, which is responsible for the detection of odorant signals by olfactory neurons. Due to the lack of phylogenetic signal present in the sequences of this multigene family, which evolves very rapidly, phylogenetic analyses have so far never made it possible to precisely retrace how ORs diversified prior to the appearance of present-day insect orders, and what the drivers of this diversification were. For example, one may suspect that the adaptation of ORs to odors emitted by plants constituted a critical step in insect evolution during the "angiosperm terrestrial revolution," which occurred at the end of the Cretaceous, but nothing currently allows this to be asserted.
There are very nice examples, notably in drosophilids, derived from comparisons between closely related species and documenting mechanisms of OR adaptation to certain signals. However, what the authors attempt to do in this work is to produce a macroevolutionary analysis at the scale of insects as a whole, based almost exclusively on bioinformatic analyses. To do this, they annotated OR genes in about one hundred insect species and developed pipelines for analyzing sequence similarity, structural similarity and functional similarity, the latter being estimated through a molecular docking approach. An important element in the evolution of insect ORs is the appearance of a unique co-receptor, called Orco, which appears to be an OR that has lost the ability to bind odorants. In addition to the large-scale bioinformatic analysis, the authors also aim to explore more specifically the factors that favored the emergence of Orco and the selective advantage conferred by the existence of OR-Orco complexes.
Given the importance of odorant receptors in insect biology and in their adaptation to different environments and lifestyles, retracing their evolutionary history is indeed a major question in evolutionary biology. In principle, this type of work therefore has the potential to become a reference in the field and to provide a basis for significant scientific advances.
Major strengths and weaknesses
The sampling chosen for collecting OR sequences is very impressive, with more than 100 insect families represented, covering most of the major orders. This sampling appears appropriate for the question being addressed. The analysis pipeline used to collect the sequences makes sense, relying on homology-based annotation tools coupled with a structure-based filter. Nevertheless, one can note aberrant numbers of ORs for certain species (much lower than reality). A lower number of OR genes is somewhat expected, as the authors chose to apply a fairly stringent filter on sequence quality (based on predicted 3D structure), which reduces the number from 14,000 to 9,000. This choice seems logical given the subsequent use of these data, but it inevitably leads to data loss. However, the low number of genes also results from the fact that the pipeline did not function correctly for all genomes.
In the revised version of the manuscript, the authors included a benchmarking step, which is a good point. They compared their OR gene annotations with previous reports in the same species. Unfortunately, this comparison is irrelevant because the chosen reference OR repertoires are actually a mix of transcriptome and genome annotations. Furthermore, comparisons with entire OR gene repertoires essentially demonstrate that their annotation was of good quality for species in which OR sequences were already present in the query OR database, but poor for species in which they were not. Therefore, these supplementary analyses made by the authors are not particularly in favor of an overall good quality of OR gene repertoires in the >110 species studied. The fact that some OR genes may be missing and that the total number may not be exact for each species is not prohibitive for studying the evolution of the family on a broad scale. However, it does call into question the correlation between the number of ORs and lifestyles and diets.
From the dataset collected, the authors attempted to categorize ORs in several ways, starting with the reconstruction of sequence similarity networks. The approach is interesting, but fails to reveal homology relationships between ORs from species belonging to different insect orders. So it is unclear what the advantage of this approach is compared with the "classical" phylogenetic approach.
The clustering based on structure also leads to the identification of a majority of "order-specific" clusters, which does not provide major insights into the evolution of ORs. However, the authors highlight a group of ORs in flies that appear to possess an unusual intracellular region, as well as a cluster of OR shared across many insect orders that exhibit a larger binding cavity. This is really interesting, although more relevant to OR structure than to their evolution.
The analysis of structural diversity then leads the authors to focus on the Orco co-receptors, which are characterized by modifications of the binding pocket and the appearance of an extracellular loop that could explain the loss of the ability to bind odorant molecules. This part, which relies on in vitro experiments, is interesting and constitutes the most striking result of this study, which could in itself have been the subject of a separate manuscript.
The rest of the manuscript is based on the prediction of OR response spectra using molecular docking. The work that has been carried out is extremely substantial, and the objective of linking clusters based on sequence similarity or 3D structural similarity with functional categories is entirely relevant. The docking score threshold used was chosen thoughtfully, which is very good, and according to the calculation performed should ensure a true positive rate of more than 20%, which is excellent in such a docking analysis. But in the absence of functional validation, this 20% true positive rate is not sufficient to extrapolate OR function, and docking-derived binding breadth measures used in the remaining of the manuscript have to be taken with caution, as acknowledged by the authors themselves. Consequently, the chance that results of this part of the work will enlighten the evolution of OR on a broad scale is low. For example, the fact that insect lineages that emerged after the Permian-Triassic extinction have more broadly-tuned OR is an interesting observation, yet remains highly speculative.
In summary, despite the large number of analyses performed, the authors do not really succeed in achieving the stated objective of reconstructing the evolutionary history of insect ORs, and the results obtained do not strongly support all the conclusions regarding the links between OR repertoires and environment or lifestyle.
-
Reviewer #2 (Public review):
The remarkable evolvability of the olfactory system enables animals to rapidly adapt to dynamic and chemically complex environments. Over the past two decades, substantial effort has been devoted to uncovering the evolutionary principles that drive the diversification of odorant receptors (ORs), yielding key insights into the forces shaping their striking variability in both vertebrates and insects. In this manuscript, Zhang and colleagues analyze the OR repertoires of over 100 insect species, leveraging sequence and structural similarity to infer patterns of gene family evolution within this diverse and ecologically important clade. By integrating sequence-based and structure-based comparisons, their study builds on a compelling and recently emerging line of research made possible by the advent of …
Reviewer #2 (Public review):
The remarkable evolvability of the olfactory system enables animals to rapidly adapt to dynamic and chemically complex environments. Over the past two decades, substantial effort has been devoted to uncovering the evolutionary principles that drive the diversification of odorant receptors (ORs), yielding key insights into the forces shaping their striking variability in both vertebrates and insects. In this manuscript, Zhang and colleagues analyze the OR repertoires of over 100 insect species, leveraging sequence and structural similarity to infer patterns of gene family evolution within this diverse and ecologically important clade. By integrating sequence-based and structure-based comparisons, their study builds on a compelling and recently emerging line of research made possible by the advent of AlphaFold, which has previously clarified the phylogenetic relationship between insect Ors and the gustatory receptor gene family and revealed the unexpectedly deep evolutionary origins of this ancient structural fold.
Applying this approach to a large set of ORs derived from species throughout the insect phylogeny, the authors confirm many previously reported patterns of OR evolution. More importantly, with their large-scale approach the authors found new structural features of different Or clades, resulting in intriguing testable hypotheses about Or evolution that have the potential to support future directions in studying this important gene family. For example, the authors identify a structural feature mostly unique to the OR co-receptor ORco, a beta-sheet in EL2, which they functionally show reduces odorant binding affinity - a key aspect of ORco, which does not bind ligands in the ancestral ligand-biding site. This is a particularly strong part of the manuscript, since the authors support their in silico-derived hypothesis with functional data.
In an attempt to assess the relationship between sequence identity and structure on one hand and function on the other, the authors perform an in-silico structure prediction and chemical docking analysis. While not suitable to assess the actual function of individual receptors without functional testing, this strategy revealed potential receptor clades with differences in the number and type of docked chemicals. As a response to the previous reviews, the authors now changed their writing to refer to these results as 'docking-derived function' instead of just 'function'. While I appreciate that the authors changed their wording, this semantic change still can be read as referring to docking-derived predictions as conclusions of actual biological function. However, this is not the case. Any functional claims based solely on the docking approach need to be interpreted with caution, as in silico-only approaches are currently not well enough predictors for actual receptor function. Accordingly, I strongly recommend to remove the parts that connect receptor function to structure or ecology.
Assessing their docking analyses with respect to species ecology, the authors identify correlations suggesting that Or repertoires might follow predictable evolutionary trajectories broadly aligned with species ecology. These are interesting hypotheses. However, due to the inability to predict actual receptor function the author's claims about ecology-function relationships are not well supported. Accordingly, the authors should change the respective parts of the manuscript in way that it is clear these are currently untested hypotheses. This includes adjusting the title, since the work does not find any experimental support for ecological tuning.
I believe that future studies combining functional testing with an explicit evo-eco framework could benefit from this work and the explicit hypotheses it generates. But the authors should be careful to not overstate their in-silico derived results.
-
Author response:
The following is the authors’ response to the original reviews.
Public Reviews:
Reviewer #1 (Public review):
Objectives of the study and impact of the work:
The authors of this article primarily aim to reconstruct the evolutionary history of the insect odorant receptor (OR) family, which is responsible for the detection of odorant signals by olfactory neurons. Due to the lack of phylogenetic signal present in the sequences of this multigene family, which evolves very rapidly, phylogenetic analyses have so far never made it possible to precisely retrace how ORs diversified prior to the appearance of present-day insect orders, and what the drivers of this diversification were. For example, one may suspect that the adaptation of ORs to odors emitted by plants constituted a critical step in insect evolution during …
Author response:
The following is the authors’ response to the original reviews.
Public Reviews:
Reviewer #1 (Public review):
Objectives of the study and impact of the work:
The authors of this article primarily aim to reconstruct the evolutionary history of the insect odorant receptor (OR) family, which is responsible for the detection of odorant signals by olfactory neurons. Due to the lack of phylogenetic signal present in the sequences of this multigene family, which evolves very rapidly, phylogenetic analyses have so far never made it possible to precisely retrace how ORs diversified prior to the appearance of present-day insect orders, and what the drivers of this diversification were. For example, one may suspect that the adaptation of ORs to odors emitted by plants constituted a critical step in insect evolution during the "angiosperm terrestrial revolution," which occurred at the end of the Cretaceous, but nothing currently allows this to be asserted.
There are very nice examples, notably in Drosophilids, derived from comparisons between closely related species and documenting mechanisms of OR adaptation to certain signals. However, what the authors attempt to do in this work is to produce a macroevolutionary analysis at the scale of insects as a whole, based almost exclusively on bioinformatic analyses. To do this, they annotated OR genes in about one hundred insect species and developed pipelines for analyzing sequence similarity, structural similarity, and functional similarity, the latter being estimated through a molecular docking approach. An important feature in the evolution of insect ORs is the emergence of a unique co-receptor, called Orco, which appears to be an OR that has lost the ability to bind odorants. In addition to the largescale bioinformatic analysis, the authors also aim to explore more specifically the factors that favored the emergence of Orco and the selective advantage conferred by the existence of OR-Orco complexes.
Given the importance of odorant receptors in insect biology and in their adaptation to different environments and lifestyles, retracing their evolutionary history is indeed a major question in evolutionary biology. In principle, this type of work therefore has the potential to become a reference in the field and to provide a basis for significant scientific advances.
Major strengths and weaknesses:
The sampling chosen for collecting OR sequences is very impressive, with more than 100 insect families represented, covering most of the major orders. This sampling appears appropriate for the question being addressed. The analysis pipeline used to collect the sequences makes sense, relying on homology-based annotation tools coupled with a structure-based filter. Nevertheless, one can note aberrant numbers of ORs for certain species (much lower than reality), which indicates that the pipeline probably did not function correctly for all genomes. In the absence of a validation step comparing the results with already known OR repertoires, it is difficult to estimate the overall quality of the data. The authors chose to apply a fairly stringent filter on sequence quality (based on predicted 3D structure), which reduces the number from 14,000 to 9,000. This choice seems logical given the subsequent use of these data, but it inevitably leads to data loss. The fact that some OR genes may be missing and that the total number may not be exact for each species is not prohibitive for studying the evolution of the family at a broad scale; however, it calls into question certain results that rely on this total number, such as the correlation between the number of ORs and genome size, lifestyle, and diet.
We thank the reviewer for raising this important concern. To objectively evaluate how much our strict structural filtering may have reduced OR counts, we collected published OR annotations for species included in our study and compared those values with our structurally intact OR counts (Author response table 1). For most previously annotated species, the difference was within approximately ten ORs, indicating that our counts are generally comparable to published annotations after structural filtering. Diabrotica virgifera virgifera was a clear exception. Previous work, using transcriptomic evidence and manual annotation, reported 193 ORs, including 124 complete ORs and three pseudogenes, whereas our original pipeline retained only nine structurally intact ORs. We found that this discrepancy was mainly caused by insufficient query representation. The previous D. virgifera virgifera annotation was published in 2025, whereas our original annotation was performed in 2022, so our query set likely did not adequately represent this lineage. When we re-annotated this species using new queries, 78 complete ORs were retained after structural filtering. We have updated the D. virgifera virgifera annotation in the revised dataset.
We also considered genome size as a possible contributor to low automated recovery. Among the 115 species analyzed, D. virgifera virgifera has the third-largest genome, after Locusta migratoria and Thermobia domestica. Large genomes contain many repetitive regions, which can make automated OR annotation more difficult and may reduce the number of recovered intact gene models. The two larger-genome species in our dataset had previously been manually annotated with transcriptomic support, suggesting that automated annotation alone may be less reliable for large, repeat-rich genomes. At the same time, published OR annotations are not always free from error. For example, Propsilocerus akamusi was previously reported to have 17 ORs, whereas our pipeline retained ten complete ORs. After downloading and modeling the 17 published sequences, we found that PaOR1, PaOR5, PaOR6, PaOR9, and PaOR14 differed substantially from canonical OR structures or appeared to be incorrectly annotated (Author response image 1). Truncated genes such as PaOR4, which lacks part of the intramembrane region of TM2-4 but still clearly resembles an OR, would be retained by our strategy; sequences with more severe structural inconsistency would be excluded. Because many previous OR annotations have not been experimentally validated, it is difficult to treat all published OR counts as exact ground truth.
To test whether possible OR loss affected OR-count-based conclusions, we first removed all potential GR sequences from the dataset. For species with published OR counts, we then repeated the pGLS analyses using the larger of our structurally intact OR count and the published OR count (Fig. S1b-e). The main ecological associations were retained: OR count remained associated with larval diet, adult diet, and habitat, and was not associated with circadian rhythm. This indicates that the main OR count ecological analyses are reasonably robust to moderate underestimation caused by missed ORs or strict structural filtering.
In contrast, the association between OR count and genome size was no longer supported in this sensitivity analysis (Author response image 2a and b). We therefore removed the analysis and discussion of a negative relationship between genome size and OR count. The original pattern likely reflected the difficulty of annotating ORs in large, repeat-rich genomes rather than a reliable biological relationship.
The following revision has been added to the revised manuscript (lines 122-126): " We then repeated the phylogenetic generalized least squares (pGLS) analyses using the larger of our structurally intact OR count and the published OR count, and the main ecological associations were retained. This indicates that the main OR-count ecological analyses are reasonably robust to moderate underestimation caused by missed ORs or strict structural filtering."
These new benchmark and sensitivity analyses directly strengthen the evidence supporting our dataset and its ecological interpretations.
From the dataset collected, the authors attempted to categorize ORs in several ways, starting with the reconstruction of sequence similarity networks. The approach is interesting, but in the end, the results do not seem to be sufficiently exploited, and it is not obvious what the advantage of this approach is compared with the "classical" phylogenetic approach, which generally fails to reveal homology relationships between ORs from species belonging to different insect orders. Here again, the majority of the clusters identified are "order-specific," and when this is not the case, the authors did not attempt to exploit the results. For example, clusters SeqC26 or SeqC28, which appear to be shared by many insects, are potentially very interesting. It might have been relevant to combine this similarity-based clustering approach with phylogenetic reconstructions within each shared cluster.
We thank the reviewer for this insightful suggestion. We agree that the previous version did not sufficiently explore the evolutionary information contained in shared sequence communities such as SeqC26 and SeqC28. Following the suggestion, we extracted the sequences from these key shared communities and reconstructed within-community gene trees to search for orthologous groups.
Because shared SeqCs may contain conserved genes across insect orders, we used the gene trees of SeqC26 and SeqC28 to identify candidate cross-order orthologous groups. In SeqC26, we identified 56 orthologous groups, 11 of which included more than ten species. Among these 11 larger groups, 4 spanned multiple insect orders. One example, OGG54, includes sequences from Orthoptera, Blattodea, Trichoptera, and Hymenoptera; after excluding the possibility of GR contamination, this group includes locust LmigOR5 and LmigOR4. LmigOR5 has been reported to bind geranyl acetone and to be associated with avoidance behavior in the migratory locust (Chang et al. 2023). The functions of the corresponding receptors in other insect orders remain to be tested, and we now present these as candidate conserved modules rather than confirmed functional orthologs. In contrast, we did not identify clear cross-order orthologous groups in SeqC28. This suggests that the ORs in SeqC28 are more similar at the sequence-community level, but do not necessarily reflect traceable orthologous relationships.
The following revision has been added to the revised manuscript (lines 156-164): "SeqC26 and SeqC28 are large shared clusters that may contain cross-order orthologous relationships. In SeqC26, we identified 56 orthologous groups, four of which spanned multiple insect orders. OGG54 included ORs from Orthoptera, Blattodea, Trichoptera, and Hymenoptera. This group contains the locust receptors LmigOR5 and LmigOR4. LmigOR5 has been reported to bind geranyl acetone and to be associated with avoidance behavior in the migratory locust. The functions of the corresponding receptors in other insect orders remain to be tested, and we now present these as candidate conserved modules rather than confirmed functional orthologs. In contrast, we did not identify clear cross-order orthologous groups in SeqC28."
The clustering based on structure also leads to the identification of a majority of "orderspecific" clusters, but once again, the clusters shared by several orders are not truly exploited, which does not provide new insight into the evolution of ORs. However, the authors highlight a group of ORs in flies that appear to possess an unusual intracellular region. This is interesting, although it is a result more relevant to OR structure than to their evolution. The function of these ORs in Drosophila melanogaster, if it is known, is not discussed.
We thank the reviewer for this useful suggestion. Similar to the sequence communities, the structural communities also contain broadly shared groups. In the revised analysis, we focus in particular on StrC23, the only structural community present in 12 insect orders. StrC23 accounts for 46.3% of all structurally intact ORs in our dataset (4124 of 8905 ORs).
Given the extremely low sequence similarity among insect ORs, the existence of such a broad conserved structural community is important. We found that StrC23 has a significantly larger binding-pocket volume than other ORs and Orco (Fig. S3 d). From a structural perspective, this suggests that StrC23 receptors may have greater potential to accommodate diverse VOCs. We therefore interpret StrC23 not as sequence-level conservation, but as conservation at the level of OR structural evolution: many species appear to retain a large set of ORs with relatively large binding pockets, which may provide a structural basis for broad docking-derived VOC binding potential before lineage-specific OR diversification occurs.
Following the reviewer’s suggestion, we also examined available functional data for the long-IL3 receptors in StrC17. The long-IL3 orthologous group includes Drosophila melanogaster DmOr13a (droMel44 in our dataset) and Bactrocera dorsalis BdorOR13a (bacDor_18 in our dataset). Previous studies indicate that both receptors respond to 1-octen-3-ol (Kreher et al. 2008; Xu et al. 2023). Earlier work also identified a group of Dipteran fly OR homologs, including DmOr13a and BdorOR13a, as 1octen-3-ol-specific responsive receptors associated with oviposition behavior(Liu et al. 2023). When we modeled these homologous receptors, they also showed an extended IL3 region.
We therefore propose that the long-IL3 ORs identified here likely correspond to the previously reported fly homologs involved in 1-octen-3-ol responses. However, 1-octen-3-ol has different behavioral functions in different species: it acts as an attractant in blood-feeding mosquitoes and tsetse flies (Hall et al. 1984; Kline et al. 2007), contributes to plant-host localization in parasitoids(Morawo and Fadamiro 2016), functions as an aggregation cue in some beetles(Pierce et al. 1989), and is often associated with oviposition regulation in flies (Kreher et al. 2008; Liu et al. 2023). Mosquito receptors known to detect 1-octen-3-ol, such as AgOR8, AaOR8, TaOR8, and CquiOR118b(Lu et al. 2007; Xu et al. 2015; Dekel et al. 2016; Frunze et al. 2024), do not contain a long IL3 region (Fig. S3 e). This raises the possibility that the long IL3 structure in fly homologs may relate to the oviposition-associated role of 1-octen-3-ol in flies.
Because OR ligand binding is primarily mediated by the binding pocket, IL3 may not directly determine ligand binding. Instead, it may influence downstream regulation of olfactory responses. Mutational analysis of Orco suggests that IL3 can regulate channel activation and Orco-dependent olfactory responses, indicating that variation in intracellular loops may affect response efficiency or sensitivity(Turner et al. 2014; Bobkov et al. 2021). We now discuss this only as a plausible mechanism requiring future experimental validation.
The following revision has been added to the revised manuscript (lines 196-202): "StrC23 is the only structural community present in 12 insect orders and accounts for 46.3% of all structurally intact ORs in the dataset. We found that StrC23 has a much larger binding-pocket volume than other ORs and Orco. From a structural perspective, this suggests that StrC23 receptors may have greater potential to accommodate diverse VOCs. This reflects conservation at the level of OR structural evolution: many species appear to retain a relatively large set of ORs with larger binding pockets, which may provide a structural basis for structure-derived volatile organic compound binding potential before species-specific OR diversification."
The following revision has also been added (lines 408-419): "The long-IL3 OR orthologous group includes Drosophila melanogaster DmOr13a and Bactrocera dorsalis BdorOR13a, both of which respond to 1-octen-3-ol and belong to a group of dipteran fly OR homologs associated with oviposition behavior. Modeling of these homologous ORs showed that they also contain a long IL3 region. We therefore propose that long-IL3 ORs correspond to these fly homologs that recognize 1-octen-3-ol. However, 1-octen-3-ol has different behavioral functions in different species: it acts as an attractant in blood-feeding mosquitoes and tsetse flies, whereas in flies it is usually associated with oviposition regulation. Mosquito receptors known to detect 1-octen-3-ol, such as AgOR8, AaOR8, TaOR8, and CquiOR118b, do not contain a long IL3 region. This suggests that the long IL3 structure in fly homologs may be related to the oviposition-associated role of 1-octen-3-ol in flies, which requires future experimental validation."
The analysis of structural diversity then leads the authors to focus on the Orco co-receptors, which are characterized by modifications of the binding pocket and the emergence of an extracellular loop that could explain the loss of the ability to bind odorant molecules. This part, which relies on in vitro experiments, is interesting and constitutes the most striking result of the study, which could in itself have been the subject of a separate manuscript. However, the molecular dynamics modelling does not add anything in the way it is conducted (5 ns is too short).
We thank the reviewer for the positive assessment of our Orco results and for raising this important concern. We agree that 5 ns is insufficient to evaluate the conformational stability or long‑term dynamics of the receptor complex. However, our simulations were designed specifically to examine whether the EL2 β‑sheet could sterically affect the early movement of VOCs near the extracellular entrance of the binding pocket. Importantly, rather than relying on a small number of long trajectories, we performed 60 independent short simulations with the explicit goal of capturing a statistical trend in how VOCs respond to this local steric constraint. Within 5 ns, VOC trajectories were visibly altered by the EL2 β‑sheet. Across the 60 replicates, VOCs reached the binding pocket less frequently in Orco proteins containing this structure, revealing a consistent trend (Fig. 4C and Movie S1). We therefore consider this timescale sufficient to detect the early steric response of small molecules, and the large number of repeats provides statistical confidence in this conclusion.
The following revision has also been added (lines 688–691): “It should be noted that the molecular dynamics simulations were designed to examine the early trajectories of VOCs near the extracellular entrance of the binding pocket, rather than to assess the conformational stability of the receptor complex. A duration of 5 ns is sufficient for small molecules to respond to local steric constraints.”
The rest of the manuscript is based on the prediction of OR response spectra using molecular docking. The work that has been carried out is extremely substantial, and the objective of linking clusters based on sequence similarity or 3D structural similarity with functional categories is entirely relevant. Nevertheless, I see two major problems with this in silico functional analysis:
(1) The docking score threshold used was chosen thoughtfully, which is very good, and according to the calculation performed, should ensure a true positive rate of more than 20%, which is excellent in such a docking analysis. But in the absence of functional validation, this 20% true positive rate is not sufficient to extrapolate OR function as the authors do in the remainder of the manuscript. The risk of error remains too high to compare in such detail the function of ORs from insects with different lifestyles or diets.
We thank the reviewer for pointing out this important limitation of our computational functional interpretation. We fully agree that, although the threshold calibrated from available experimental data improves enrichment, a true-positive rate of approximately 20% is not sufficient to make deterministic claims about individual OR-VOC pairs. We therefore do not equate docking predictions with true OR response spectra. To prevent any misinterpretation, we have undertaken a major revision of our terminology and framing throughout the manuscript:
The main purpose of docking in our study is not to predict the real ligand of every OR or to compare the exact functional properties of individual receptors across lifestyles. Because the study includes 115 insect species, thousands of ORs, and a large VOC set, systematic experimental validation of all ORVOC combinations is currently not feasible. Instead, we use docking as a high-throughput and internally comparable theoretical screening method to estimate the potential binding tendencies of OR repertoires toward different functional-group VOCs.
Under this framework, we focus on repertoire-level trends generated by a uniform structural modeling, docking, and scoring pipeline. The value of this analysis is to provide candidate receptors, VOC categories, and ecological associations for future experimental testing. In the revised manuscript, when the results are based solely on docking analyses, we avoid directly referring to them as OR functions. Instead, we add the prefix “docking-derived” to prevent potential misunderstanding. We also explicitly state that confirming specific OR ligands and behavioral functions will require heterologous expression, electrophysiology, calcium imaging, genetic perturbation, or behavioral assays.
The following limitation statement has been added to the revised manuscript (lines 285-288): "Although this threshold enriches experimentally responsive OR-VOC pairs, the resulting hit rate is not sufficient to support deterministic inference for individual receptor-ligand pairs. Therefore, subsequent analyses were interpreted at the OR repertoire level rather than as experimentally validated OR response spectra.”; (lines 292-293) “It should be noted that dFunCs represent docking-derived functional communities rather than experimentally confirmed functional classes.”
(2) The six functional clusters identified are only slightly different from one another, with similar detection of all chemical families except acids and amines (which was expected, given that these families are a priori detected by IRs rather than ORs). This shows that even though the approach is relevant and deserves to be tested, it cannot be used to establish a link between groups/lineages of ORs and response spectra at the scale of insects as a whole. This is reflected in the final analysis by the fact that there is no visible link between sequence or structural clusters and functional clusters. Given the uncertainty surrounding the docking results, the entire subsequent analysis of the relationship between the Binding Breadth Index and ecological variables is highly questionable.
We thank the reviewer for this key comment. We agree that the differences among dFunCs for any single VOC functional group are often modest. However, the dFunC classification was not based only on whether an OR strongly recognizes one specific functional-group category. Rather, it was based on each OR’s overall docking-derived binding profile across a large VOC set.
In other words, the differences among dFunCs mainly reflect combinations of relative binding tendencies across multiple VOC categories, rather than a strong difference for one chemical family alone. Therefore, even if the predicted binding level for a single functional group differs only slightly among dFunCs, their multivariate profiles can still form distinct dFunCs.
We also agree that SeqCs, StrCs, and dFunCs do not show a simple one-to-one correspondence. This is expected because the three classifications are based on different information: sequence similarity, overall structural similarity, and docking-derived binding-profile similarity. At present, there is no direct evidence that insect OR sequence, structure, and potential binding profile maintain a conserved one-to-one relationship across the insect class. We also cannot exclude the possibility that false positives in docking reduce the resolution of the functional classification.
To avoid overinterpretation, we now refer to FunCs as docking-derived functional communities (dFunC) and define BBI as docking-derived BBI. This metric is intended for macro-scale relative comparisons, not for assigning true ligands or specific ecological functions to individual ORs. Ecological interpretations based on dFunC or BBI have been substantially toned down and are presented as candidate trends for future functional validation.
The following limitation statement has been added to the revised manuscript: "Ecological associations based on BBI should be interpreted as repertoire-level predicted binding trends, not as direct evidence for lifestyle-specific OR function or behavioral olfactory capacity."
This reframing ensures our conclusions are appropriately supported by the computational evidence and aligns the manuscript's claims with its methodological strengths.
Finally, the evolutionary analysis proposed to conclude that the work suffers from an incorrect interpretation: ORs of non-holometabolous insects cannot be considered equivalent to those of species that existed before the Permian-Triassic extinction. The fact that a locust or a cockroach has more narrowly tuned ORs than holometabolous insects does not mean that this was also the case for ancestral insects. To advance this type of conclusion, it would be necessary to conduct a phylogenetic analysis and reconstruct ancestral states, which is not the case here.
In summary, despite the large number of analyses performed, the authors do not succeed in achieving the stated objective of reconstructing the evolutionary history of insect ORs, and the results obtained do not sufficiently support the conclusions regarding the links between OR repertoires and environment or lifestyle.
We thank the reviewer for this important point. We agree that a rigorous test of changes in ancestral OR functional spectra before and after the EPME would require ancestral reconstruction within a phylogenetic framework based on reliable orthologous groups. For insect ORs, this is currently limited by rapid sequence divergence, very restricted cross-order and even within-order orthology, and frequent gene duplication, loss, and lineage-specific diversification.
Therefore, our current data cannot reliably reconstruct complete ancestral OR repertoires for different insect-order nodes, nor can they accurately trace extant ORs back to ancestral OR states before and after the EPME. The analysis is better understood as a comparison of OR repertoire composition among extant insect lineages, rather than as a comparison of ancestral states.
If major environmental transitions influenced the long-term evolution of insect ORs, such effects may appear either as traceable orthologous changes or as broad differences in extant repertoire composition and binding-profile composition. However, differences observed among extant lineages cannot be directly attributed to the EPME because they may also reflect later lineage-specific expansions, gene losses, and ecological adaptations.
Accordingly, we have revised the conclusion. We no longer state that ancestral insect OR functional spectra changed before and after the EPME. Instead, we state that comparisons among extant insect repertoires reveal differences in docking-derived binding-profile composition among lineages. These patterns may provide hypotheses for exploring links between major geological/ecological transitions and long-term olfactory evolution, but they require broader phylogenetic sampling, ancestral reconstruction, and functional validation.
The following revision has been added to the revised manuscript (lines 427-430): "We also detected differences in dFunC composition between extant lineages whose order-level origins fall before and after the EPME. Lineages originating after the EPME showed a higher proportion of broad-tuned ORs, likely influenced by multiple factors."
We also added the following clarification (lines 444-450): “However, because insect ORs evolve rapidly and cross-order orthologous relationships are limited, the current dataset does not allow reliable reconstruction of complete ancestral OR repertoires at deep insect nodes. The EPME-related comparison should therefore be interpreted as a descriptive comparison among extant lineages, not as a direct test of ancestral OR functional changes before and after the EPME. This hypothesis will require further testing through broader taxon sampling, reliable orthology assignment, ancestral-state reconstruction, and functional assays.”
This revision directly addresses the reviewer's concern by removing the unsupported causal claim and reframing the analysis within the appropriate, evidence-based scope of our study, thereby strengthening the manuscript's contribution as a source of robust comparative patterns and testable macroevolutionary hypotheses.
Reviewer #2 (Public review):
The remarkable evolvability of the olfactory system enables animals to rapidly adapt to dynamic and chemically complex environments. Over the past two decades, substantial effort has been devoted to uncovering the evolutionary principles that drive the diversification of odorant receptors (ORs), yielding key insights into the forces shaping their striking variability in both vertebrates and insects. In this manuscript, Zhang and colleagues analyze the OR repertoires of over 100 insect species, leveraging sequence and structural similarity to infer patterns of gene family evolution within this diverse and ecologically important clade. By integrating sequence-based and structure-based comparisons, their study builds on a compelling and recently emerging line of research made possible by the advent of AlphaFold, which has previously clarified the phylogenetic relationship between insect Ors and the gustatory receptor gene family and revealed the unexpectedly deep evolutionary origins of this ancient structural fold.
Applying this approach to a large set of ORs derived from species throughout the insect phylogeny, the authors confirm many previously reported patterns of OR evolution. Unfortunately, the way these results are presented lacks clarity in what is already known from previous work in the field versus what is a novel finding based on the analysis of this dataset.
We thank the reviewer for pointing this out. We agree that the original manuscript did not distinguish previously established findings from the novel results of the present study with sufficient clarity. Rapid OR evolution, lineage-specific expansion, and associations between OR repertoires and ecological traits have been reported in several insect groups. We have revised the manuscript to acknowledge these studies more explicitly and to indicate which results confirm known patterns, which extend them across a broader phylogenetic scale, and which arise from our new analyses.
Our study contributes more than a confirmation of previous observations. To our knowledge, it provides the first integrated analysis of OR evolution across 115 insect species that combines sequence similarity, structural conservation, and docking-derived binding profiles within a unified macroevolutionary framework. This analysis identifies class-wide repertoire patterns and previously unrecognized structural features, including the conserved structural cluster StrC23, that could not be evaluated using narrower taxonomic datasets.
The study also provides a mechanistic insight into Orco specialization. We experimentally demonstrate that the distinctive EL2 β-sheet of Orco reduces ligand binding affinity, supporting its contribution to the reduced odorant binding capacity of this conserved coreceptor. In addition, because exhaustive experimental characterization of the large insect OR family is currently impractical, our experimentally calibrated computational pipeline provides a framework for identifying repertoire-level trends, generating testable hypotheses, and prioritizing receptors for future functional studies.
The reviewer raises several specific examples of the distinction between previous knowledge and novel findings in the comments below. We address each of these points in the corresponding responses. We believe these revisions more clearly position our contribution as testing, extending, and integrating previously reported patterns while also identifying new class-wide structural and functional features.
It is unclear how complete the odorant receptor sets are. I recommend benchmarking the pipeline by comparing its output to a gold standard and a frequently vetted complete OR set, such as that of Robertson and Wanner 2006 or similar.
We thank the reviewer for this suggestion. We agree that without benchmarking, readers cannot adequately evaluate the completeness and accuracy of the OR annotation workflow. Following the suggestion, we performed detailed comparisons using three relatively well-curated repertoires: Drosophila melanogaster, Anopheles gambiae, and Bombyx mori. Because the species included in our study did not include Apis mellifera from Robertson and Wanner 2006, we did not use that dataset as the benchmark.
The benchmark addresses three questions. First, how well does the automated genome-based OR annotation workflow recover full-length ORs reported in previous studies? Second, does the structural filtering step improve the structural quality and reliability of the OR dataset? Third, for candidates in tandem-repeat regions where exon-merging errors may occur, do the candidate ORs share genomic positions with the most similar literature ORs, or are they more likely to represent distinct candidates? For Drosophila melanogaster, the accepted repertoire contains 60 OR genes and 65 protein products including splice variants. At the gene level, the automated workflow recovered most benchmark ORs before structural filtering, but Or47b and Or98b were not annotated. After structural filtering, fragmented or structurally incomplete ORs were removed, leaving 56 ORs that otherwise correspond one-to-one with the benchmark ORs. Two ORs that had been annotated before filtering, DmOr59a and DmOr85e, were removed by the structural filter because DmOr59a was fragmented and DmOr85e lacked TM7 in the predicted structural model (Author response image 3).
For Anopheles gambiae, the published repertoire includes 79 OR genes. Our automated workflow initially annotated 77 ORs, including 72 published ORs. After removing fragments and five full-length literature ORs, the final structurally filtered set contained 67 ORs. We inspected the five removed full-length ORs and found that AgOr52, AgOr47, and AgOr64 lacked a complete transmembrane region, whereas AgOr58 and AgOr6 contained all transmembrane regions but showed local deletions in some regions (Author response image 3). This indicates that structural filtering removes clearly incomplete structures, but can also be conservative enough to exclude a small number of true ORs with atypical transmembrane-region length variation.
For Bombyx mori, the published repertoire contains 66 OR genes including two pseudogenes. Our automated workflow initially annotated 103 ORs, including 60 previously reported ORs. After removing fragments and seven literature ORs, the final set contained 64 ORs, of which 53 corresponded to literature annotations. For structurally retained candidates that did not map one-to-one to literature ORs, we checked genomic locations (Table R2). Most were not located on the same chromosome or in the same tandem region as their most similar literature ORs, making exon-merging artifacts less likely. They may therefore represent previously unannotated OR candidates, although transcriptomic evidence or manual annotation will be needed for confirmation.
Overall, the benchmark shows that the automated workflow recovers most known OR repertoires and that structural filtering produces a high-confidence set suitable for structural comparison and docking. We now explicitly describe the final dataset as a set of structurally intact ORs rather than a complete OR repertoire for every species, and we discuss how strict filtering may underestimate OR counts. We added a benchmark description to the main text (lines 100-104) and included the detailed comparison strategy and results in Supplementary Text S1: "Benchmarking against existing OR repertoires from Drosophila melanogaster, Anopheles gambiae, and Bombyx mori showed that the annotation workflow provides high-confidence annotations suitable for downstream structural comparison and docking (see Supplementary Text S1, Table S1-3)."
Using their structural clustering approach, the authors identify a structural feature mostly unique to the OR co-receptor ORco, a beta-sheet in EL2, which they functionally show reduces odorant binding affinity - a key aspect of ORco, which does not bind ligands in the ancestral ligand-binding site. This is a particularly strong part of the manuscript, since the authors support their in silico-derived hypothesis with functional data.
We thank the reviewer for the positive assessment of the Orco-related work.
Lastly, in an attempt to assess the relationship between sequence identity and structure on one hand and function on the other, the authors perform an in silico structure prediction and chemical docking analysis. As it stands, this part is on the more speculative side since the docking approach has not been verified with available functional datasets.
We thank the reviewer for this comment and agree that the docking analysis is predictive. Molecular docking cannot be treated as experimental evidence for OR response spectra or as a definitive assignment of ligand specificity for individual OR-VOC pairs.
In the revised manuscript, we clarify that docking is used as a unified, comparable theoretical framework for macro-scale OR repertoire comparison. It is meant to estimate docking-derived binding potential across thousands of ORs and many VOCs, not to replace heterologous expression, electrophysiology, calcium imaging, or behavioral validation.
We also clarify that the docking framework was not used without any empirical calibration. We used existing OR-VOC experimental response data to examine the relationship between docking score and experimental hit rate, and we evaluated the ability of docking-derived labels to recover functional tendencies in available datasets. These analyses support the use of docking for repertoire-level enrichment and hypothesis generation, but they do not justify deterministic conclusions for individual receptor-ligand pairs. The revised manuscript now states this limitation explicitly.
The following revision has been added to the revised manuscript (lines 285-288): "Although this threshold enriches experimentally responsive OR-VOC pairs, the resulting hit rate is not sufficient to support deterministic inference for individual receptor-ligand pairs. Therefore, subsequent analyses were interpreted at the OR repertoire level rather than as experimentally validated OR response spectra."
Summary of Major Revisions:
In direct response to the eLife Assessment and reviewer comments, our revisions have systematically strengthened the evidence and clarified the interpretation:
(1) Added critical validations: Benchmarking of the annotation pipeline and sensitivity analyses for ecological correlations.
(2) Deepened evolutionary analysis: Phylogenetic exploration of shared clusters and structural characterization of StrC23.
(3) Clarified the scope and limitations of the computational functional analysis: Adopted “docking-derived” terminology and specified that the results represent hypothesis-generating, repertoire-level comparative trends rather than experimentally validated receptor functions.
(4) Corrected overinterpretations: Revised conclusions regarding the EPME and holometabolous/non-holometabolous comparisons to be descriptive and hypothesis-generating. '
(5) Improved scholarly accuracy: Updated framing to properly acknowledge prior work and corrected minor points throughout.
We believe that these revisions address the concerns underlying the assessment of incomplete evidence and provide stronger, more appropriately qualified support for the manuscript’s main conclusions.
Recommendations for the authors:
Reviewer #1 (Recommendations for the authors):
The annotation pipeline produces a number of genes that is slightly lower than reality, which is expected given the presence of a fairly stringent filter. However, the number is far too low for certain species, for example, Harmonia or Diabrotica. Whenever possible (for example, for Drosophila, Bombyx, Anopheles, etc.), adding a benchmarking step by comparing the results obtained with already identified repertoires would be a real improvement. Moreover, the dataset contains sequences of gustatory receptors that must be removed in order not to bias the analysis (CO₂ receptors in Drosophila melanogaster and Bombyx mori, for example, based on what I was able to verify). It might therefore be necessary to add a specific step in the pipeline to check for the presence of GRs in the data, since the sequences and 3D structures can be quite similar to those of ORs and therefore difficult to separate.
We thank the reviewer for this careful observation. To evaluate OR annotation counts, we compared our results with published OR annotations for previously annotated species. Except for Diabrotica virgifera virgifera, most species differed from published counts by fewer than ten ORs, supporting the general reliability of our annotation counts. For D. virgifera virgifera, the very low original count was likely caused by a combination of insufficient query representation and the difficulty of annotating ORs in a large genome. The previous D. virgifera virgifera annotation was published in 2025, whereas our original annotation was performed in 2022, so our query set likely did not adequately represent this lineage. When we re-annotated this species using new queries, 78 complete ORs were retained after structural filtering. We have updated the D. virgifera virgifera annotation in the revised dataset. Large-genome insects over 2 Gb are rare in our dataset, and the sensitivity analysis indicates that this issue does not substantially affect the main ecological associations based on OR count. Regarding gustatory receptor contamination, we found that a small number of sequences labeled as olfactory receptors but actually corresponding to GRs had been retained as queries because of an operational error during OR annotation. To remove potential GRs systematically, we collected 1,418 insect GR genes from the literature and databases and combined them with the OR library to create a receptor reference set. We then compared all annotated OR candidates against this reference set using BLASTP and removed candidates whose best match was a GR. In total, we identified and removed 76 potential GRs distributed across 38 species. These included the Drosophila CO2 receptor genes Gr63a and Gr21a, as well as Bombyx mori BmGr10 and BmGr9, which the reviewer specifically noted. All downstream analyses have been repeated using the corrected dataset. We are grateful for this careful observation, which substantially improved the rigor of the dataset.
Both Reviewer 1 and Reviewer 2 requested benchmarking against known OR repertoires, and we agree that this is necessary. We therefore benchmarked the workflow against Drosophila, Anopheles, and Bombyx repertoires, evaluating recovery of known ORs, the effect of structural filtering, and potential exon-merging artifacts in tandem regions. We further added sensitivity analyses using the larger of structurally intact OR counts and published OR counts for species with available literature data. These analyses are now included in the revised Results and supplementary materials.
The revised text is as follows (lines 100-104): "Benchmarking against existing OR repertoires from Drosophila melanogaster, Anopheles gambiae, and Bombyx mori showed that the annotation workflow provides high-confidence annotations suitable for downstream structural comparison and docking (see Supplementary Text S1, Table S1-3)."
For the SSN approach, the validation by comparison with the phylogeny of hymenopterans is a good idea, but the result is not as convincing as presented in the manuscript since the cluster SeqC25 does not correspond to a monophyletic group. This, therefore, raises the question of the evolutionary information that can be drawn from this clustering approach. The validation with lepidopteran PRs is also a good idea, but it would be necessary to verify exactly to which PR clades the sequences used for the analysis belong, in order to determine which clade corresponds to each of the identified SeqC clusters.
We thank the reviewer for this important point. For the Hymenoptera comparison, we agree that SeqC25 does not correspond to a strict monophyletic clade in existing Hymenoptera OR phylogenies. We have clarified that the SSN approach is not intended to replace traditional phylogenetic trees or to reconstruct ancestor-descendant relationships among OR communities.
SeqC25 should be interpreted as a sequence-similarity community that is strongly expanded in Hymenoptera and partially corresponds to previously defined Hymenoptera OR subfamilies, but not as a strict phylogenetic clade. SeqCs describe modular boundaries in OR sequence-similarity space, whereas phylogenetic trees infer branching relationships under a specific evolutionary model. For insect ORs, where sequence similarity is often very low, duplication and loss are frequent, and tree support is often limited, the two approaches need not produce identical groupings.
We now state that the evolutionary information provided by SSNs mainly concerns sequence-similarity diversification patterns: highly expanded modules, order-specific or cross-order communities, community boundaries, and differences in repertoire composition among lineages. These results complement phylogenetic analysis, but detailed relationships within close taxa or OR subfamilies still require traditional phylogenetic methods.
For the Lepidoptera pheromone receptor (PR) validation, we agree that the previous analysis mixed experimentally validated PRs and candidate PRs, making it unclear which PR branch each SeqC corresponded to. We reorganized the PR dataset by separating classic PRs from recently reported new PR branches and mapped these sequences to SeqCs using BLASTP. Classic PRs were assigned almost entirely to SeqC6 (93.9%), whereas the new PR branch mapped to SeqC35 (Fig. S2f). Both SeqC6 and SeqC35 are shared by Trichoptera and Lepidoptera, suggesting that they may correspond to distinct PR-related sequence communities.
The following revision has been added to the revised manuscript (lines 144-150): "This comparison indicates that SSN-based communities are broadly consistent with established phylogenetic subfamilies, while capturing sequence-similarity modules rather than strictly monophyletic clades. We further evaluated the SSN using curated lepidopteran pheromone receptors (PRs). To distinguish different PR lineages, experimentally validated classic PRs and the recently reported new PR lineage were analyzed separately. Classic PRs were predominantly assigned to SeqC6 (93.9%), whereas the new PR lineage was assigned to SeqC35 (fig. S2F and Data S5), supporting the hypothesis of multiple independent origins of PRs.”
(3) In Figure 3, the cluster StrC17 appears to be present in both Lepidoptera and Diptera, whereas when looking at Figure S3, it seems to be present only in Diptera. There may be an error somewhere.
We thank the reviewer for identifying this inconsistency. After rechecking the data, we confirmed that StrC17 contains 47 ORs, of which 46 are from Diptera and one is from Lepidoptera. We have corrected Figure S3. This correction does not affect the main conclusions regarding the long-IL3 ORs, because all downstream analyses were conducted using the correct StrC17 composition.
(4) The phylogeny presented in Figure S4 does not help clarify the phylogenetic context of the Orco study. The main reason is that it is not correctly rooted: the ORs of M. rhabei should have been used as an outgroup, or the two GRs of D. melanogaster that are present (perhaps by mistake) in this analysis and that clearly diverge strongly from the rest of the sequences. In any case, this phylogenetic analysis does not seem to add much compared with what was presented in the article by Thoma et al. in 2018.
We thank the reviewer for pointing this out. The original purpose of the tree was to show that early-diverging insect ORs lack clear orthologous relationships, making it difficult to reconstruct ancestral OR states before and after Orco emergence. However, because the figure contributes little to the core conclusions and could introduce confusion, we have removed it from the revised manuscript. The revised text instead directly cites previous phylogenetic studies for the early OR/Orco evolutionary background.
(5) In Figure 5A, many VOCs appear unclassified when using molecular descriptors. Have you tried classifying VOCs using molecular fingerprints instead of selected molecular descriptors? And can you explain how those 32 descriptors were chosen? Furthermore, it is surprising not to see a "terpenoids" category given the importance of these molecules in insect chemical ecology. One would expect them to cluster together in a chemical space based on molecular descriptors; where are they in Figure 5A?
We thank the reviewer for this question. We first clarify that unclassified in Figure 5A does not mean that these VOCs were excluded from the analysis. It only means that they were not assigned to one of the nine predefined major functional-group categories. All VOCs were included in molecular descriptor calculation, chemical-space visualization, and downstream docking.
The 32 molecular descriptors were not arbitrarily selected by us. They come from the optimized odorant metric proposed by Haddad et al. 2008. That study began with 1,664 Dragon molecular descriptors, represented odorants as multidimensional physicochemical vectors, and selected 32 descriptors that best explained similarity in odor-induced neural responses across multiple published datasets. Haddad et al. further showed that this optimized descriptor set performed well across different animals, recording methods, and levels of olfactory-system organization. We therefore adopted these 32 descriptors to represent odor physicochemical space.
We agree that molecular fingerprints are another useful representation, especially for capturing substructures and scaffold information. We chose molecular descriptors rather than fingerprints because the purpose of Figure 5A is to visualize VOC distribution in a continuous physicochemical odor space, and because the Haddad descriptor set was optimized using neural response data. We have added this rationale to the revised manuscript.
We also agree that terpenoids are important in insect chemical ecology. In the original classification, VOC categories were based mainly on functional groups, whereas terpenoids are defined by biosynthetic origin and carbon skeleton rather than by a single functional group. Therefore, terpenoid VOCs are distributed across several functional-group categories, such as terpene alcohols, terpene aldehydes, and terpene ketones, and are not expected to form one independent cluster in a two-dimensional odour space.
Following the reviewer’s suggestion, we added an extra annotation for common terpenoid-related compounds and mapped them onto the odor space shown in Figure 5A. The result shows that terpenoids are distributed across multiple regions and functional-group categories rather than forming a single compact cluster (Fig. S5 c). We therefore retain the major functional-group classification for the main analyses and add terpenoids as an additional annotation, while explaining why they are not treated as a main category parallel to alcohols, aldehydes, ketones, and esters.
In addition, because the downstream analyses in this study rely on large-scale docking predictions, and because docking itself contains inherent uncertainty, we chose to use relatively basic and chemically explicit major functional-group categories. This classification reduces interpretive instability that could arise from excessive subdivision of VOC classes. By contrast, treating terpenoids as an independent primary category would group together molecules with substantially different functional-group properties, thereby increasing the complexity of interpreting docking-derived binding profiles. Therefore, in the revised manuscript, we retain the major functional groups as the core VOC classification scheme and add terpenoids as an additional annotation.
The specific revision in the revised manuscript is as follows (lines 268-271): "Although terpenoids are important in insect chemical ecology, they are distributed across multiple regions of odor space. To reduce the docking instability that could result from excessive VOC subdivision, we did not treat terpenoids as a category parallel to the nine basic functional-group categories."
(6) The methodology used to calculate hit rates with respect to functional data on known ORs is not sufficiently explained. The result could be shown for the different species (drosophila, mosquito, butterfly). In addition, ORs from the locust should also have been used for this comparison.
We thank the reviewer for this suggestion. In the revised Methods, we now describe the workflow in detail. We first compiled receptor-odorant combinations with clear response relationships from published functional experiments. For each OR-VOC pair, we performed docking using the same pipeline as in the global analysis and extracted the corresponding docking score. We then labeled each pair as response or non-response according to the original experimental data. Next, we grouped pairs into docking-score intervals and calculated, within each interval, the proportion of experimentally positive pairs, which we define as the hit rate. To reduce fluctuation from random sampling, each score interval was sampled three times. Finally, following the hit-rate modeling strategy of Lyu et al. 2019, we fitted the relationship between docking score and hit rate with a Bayesian curve-fitting approach and used the posterior expectation of the parameters to draw the hit-rate curve. The threshold was determined from the docking score at which the hit rate reached a plateau.
Following the reviewer’s request, we calculated hit-rate curves separately for Drosophila melanogaster, Anopheles gambiae, Lepidoptera, and Locusta migratoria OR data (Fig. S5). Drosophila, Anopheles, and Lepidoptera showed similar trends: the hit-rate plateau was approximately 20%, and the corresponding docking-score threshold was around -8 kcal/mol. This is consistent with the original combined analysis and indicates that the threshold was not driven by a single species dataset. The locust dataset behaved differently. Its experimental positive rate was low, approximately 5.2%, and most reported locust ORs are narrowly tuned. In such a highly imbalanced dataset, docking score was less able to enrich experimental positives to the same level as in the other datasets; most score intervals had hit rates below 10%. When the locust data were combined with Drosophila, Anopheles, and Lepidoptera, the overall plateau decreased to about 18%, and the corresponding threshold shifted to about -11 kcal/mol.
We consider this difference informative because it shows that OR tuning properties and experimental positive rates can affect the relationship between docking score and hit rate. The locust result suggests that stricter thresholds may be required for systems dominated by narrowly tuned ORs and low positive rates. However, our goal is to establish a unified empirical threshold for large-scale repertoire comparison, not to optimize a separate threshold for every lineage. We therefore retain -8 kcal/mol from the Drosophila, Anopheles, and Lepidoptera datasets as the main threshold, present the locust analysis as a sensitivity test, and note that this threshold may have lower positive-enrichment ability in narrowly tuned OR systems.
The specific revision in the revised manuscript is as follows (lines 281-285): "In addition, our singletaxon hit-rate sensitivity analysis showed similar trends for Drosophila, Anopheles, and moths. Because the locust OR repertoire had an extremely low positive rate, we did not include the locust functional data in the final threshold assessment (see Supplementary Text S2 for details)."
Methods section (lines 748-755): "We first compiled published functional datasets containing explicit OR-VOC response relationships. These datasets included ORs from Drosophila melanogaster, Anopheles gambiae, Helicoverpa armigera, Spodoptera littoralis, and Locusta migratoria. For each dataset, OR-VOC combinations were assigned response or non-response labels according to the original experimental results. Each OR-VOC pair was then docked using the same structural modeling, docking, and scoring workflow used in the global OR-VOC analysis. Pairs were grouped into docking-score intervals, and the hit rate for each interval was defined as the proportion of experimentally positive pairs among all pairs in that interval. Each score interval was sampled three times."
(7) It is not clear how the ROC curves were constructed; they appear to be drawn with very few points. The methodology needs to be explained in more detail here, since this is an important point.
We thank the reviewer for this suggestion. We have added a detailed description of ROC construction. The ROC analysis was designed to evaluate whether the dFunC labels are directionally consistent with available experimental functional data.
Specifically, a positive dFunC label for a functional-group VOC indicates that ORs in that docking-derived community are predicted, relative to the all-insect OR background, to bind more VOCs from that functional group. A negative label indicates a lower predicted binding tendency. We mapped Drosophila ORs to the dFunC classification and assigned each OR a positive or negative label for each functional group according to its dFunC. We then used the Drosophila experimental functional matrix to count measured responses of each OR to different functional-group VOCs. For each functional group, we plotted ROC curves and calculated AUC values by comparing the dFunC-predicted labels with experimental response counts. We agree that the curves are based on limited data points for some functional groups. This is because comprehensive OR-VOC functional matrices are still sparse, and many insect ORs have been deorphanized using limited odor panels that do not cover many functional groups.
The specific revision in the revised manuscript is as follows (lines 794-798): "Drosophila melanogaster ORs were mapped to dFunCs to evaluate the consistency between dFunC labels and experimental response data. For each VOC functional group, ORs were assigned positive or negative labels according to their dFunC labels, and these labels were compared with response counts from the Drosophila functional matrix to generate ROC curves and AUC values."
(8) What the BBI represents is not very clear, even though the calculation method is presented in detail. This should be clarified in the main text and/or in the figure legends.
We thank the reviewer for this suggestion. We now define BBI more explicitly in the main text. BBI, or binding breadth index, is a relative repertoire-level metric calculated from docking-derived dFunC labels. A dFunC label indicates whether ORs in a docking-derived binding-profile community show higher or lower predicted binding tendency toward a functional-group VOC category relative to the all-insect OR background. When we calculate BBI for a set of ORs, we summarize the predicted binding tendencies of the dFunCs to which those ORs belong, thereby estimating the relative potential binding breadth of that OR repertoire for the VOC category.
Thus, BBI should be interpreted only as a relative docking-derived metric. When comparing two OR groups for the same VOC functional group, a higher BBI means that the group is predicted, under our docking-derived framework, to have broader potential binding breadth. It does not directly represent experimentally confirmed olfactory breadth, odor perception, or behavioral function. We have revised the terminology accordingly to docking-derived potential binding breadth.
The following revision has been added to the revised manuscript (lines 319-321): "BBI is a docking-based, repertoire-level relative metric used to summarize the potential binding breadth of a set of ORs toward a given functional-group VOC category."
(9) Insects with saprophagous larvae in your dataset are almost exclusively flies. Therefore, the conclusions concerning the saprophagous diet could just as well result from phylogenetic constraints rather than from an adaptation to the diet.
We thank the reviewer for pointing this out. We agree that, because the currently available saprophagous larval species are mainly concentrated in Diptera, this result may be influenced by lineage effects. We have therefore toned down the interpretation and present it as a candidate association. Future inclusion of more saprophagous species from non-dipteran lineages will allow this relationship to be tested more rigorously.
The following revision has been added to the revised manuscript (lines 493-496): "This interpretation should be treated with caution, as the saprophagous larval species currently available in our dataset are mainly concentrated in Diptera and may therefore reflect lineage effects. Future inclusion of saprophagous species from broader insect lineages will help test this association more rigorously."
Reviewer #2 (Recommendations for the authors):
Main feedback
(1) This study presents an interesting approach to unify the exceptionally large OR multigene family in a single macroevolutionary framework. While this is a non-trivial task, given the low sequence similarity of ORs, the inferences broadly overlap with previous findings, suggesting that the approach works. Unfortunately, the line between previously identified patterns of OR biology and evolution and new insights derived from this study is somewhat blurry throughout the text at the moment and needs to be sharpened. As it reads, the manuscript is prone to overselling the results.
A few examples:
(a) Lines 37-41 - contrary to the claim in this sentence, previous work has described the principles of OR function, structure, and evolution to an extent that would warrant concluding that the fundamental logic of insect olfaction is actually comparatively well understood.
We thank the reviewer for this comment. We agree that previous studies have already established important principles of insect OR biology, including OR-Orco complex structure, ion-channel function, ligand recognition, and rapid OR evolution. Our intention was not to imply that the basic logic of insect olfaction remains unknown, but to emphasize that an insect-class-scale framework integrating OR sequence divergence, structural variation, and docking-derived binding-profile diversity has been lacking. We have revised the Introduction to clarify this point and to better acknowledge the contribution of previous studies.
We therefore revised this sentence in the revised manuscript as follows (lines 38-42): "However, a classwide integrative framework linking OR sequence divergence, structural variation, and docking-derived binding-profile diversity is still lacking. Such a framework is needed to distinguish broadly conserved patterns from lineage-specific features and to place species-level OR diversification in a broader macroevolutionary context."
(b) Lines 47-48 - work in several systems has emphasized the "evolutionary interplay between receptor family diversification and macroecological adaptation", both in species with highly specialized ecologies (plant host specialists, slave making ants -> these are even cited in the manuscript) and in a broader macroevolutionary context (e.g. bee diet breadth, Singh et al. 2025), which does not really justify the claim that OR evolution with respect to ecology is a profound enigma. The present study indeed presents the first one combining data throughout the entire insect tree of life, which is an impressive feat. The fact that inferences drawn from smaller datasets with smaller phylogenetic breadth are supported is a useful finding. I recommend emphasizing this.
We agree that the relationship between OR repertoire evolution and ecological adaptation has been investigated in multiple insect systems and should not be described as an unresolved “profound enigma.” The novelty of our study is to test, integrate, and extend these previously reported patterns across 115 insect species under a unified sequence-structure-docking profile framework. We have revised the text to emphasize this class-wide extension rather than overstating the unknowns in the field. The following revision has been added to the revised manuscript (lines 46-47): "Although several studies have linked OR repertoire evolution to ecological adaptation in specific insect lineages, a unified class-wide comparison remains lacking."
(c) Lines 347 - 348 - what is meant by the statement that the manuscript 'revealed the mechanism of insect olfactory perception underlying macroenvironmental influences' is unclear to me. What mechanism is referred to here? What do the authors mean by 'macroenvironmental influences'? It reads as if the authors claim to have shown how olfactory perception works in insects, which is not an accurate statement.
We thank the reviewer for pointing out that this statement was unclear. We did not intend to claim that our study reveals the mechanism of insect olfactory perception itself. Rather, our results identify class-scale associations between OR repertoire variation, ecological traits, and docking-derived binding profile composition. We have revised the Discussion.
In the revised manuscript, this sentence has been changed to the following more accurate wording (lines 399-402): "Our results reveal class-level associations between OR repertoire variation and ecological traits and identify differences in docking-derived binding-profile composition among extant insect lineages that may be relevant to long-term macroevolutionary transitions."
(d) Lines 348-350: I do not agree with the statement that the authors "describe the evolutionary process through which Orco emerged". The data that the beta-sheet in EL2 impedes ligand binding in non-orco ORs is compelling and suggests an intriguing contribution to why Orco lack ligand-binding function, but is not sufficient to comprehensively describe the evolutionary process of Orco evolution from ancestral ORs. This would require more sequences of early-branching insect species. Accordingly, this statement should be toned down.
We agree that the original wording overstated the extent to which our data describe Orco emergence. Our results identify structural features, including the EL2 beta-sheet and specialized binding-pocket properties, that may have contributed to the loss of ligand-binding function during Orco specialization. However, a complete reconstruction of Orco origin will require broader sampling of early-diverging insect lineages and more reliable ancestral-state reconstruction. We have revised the relevant statements accordingly.
We therefore toned down this statement in the revised manuscript and described it more accurately as follows (lines 402-405): "We further examined Orco specialization within the insect 'conserved chassisdiversified sensors' model and identified structural features, including the EL2 beta-sheet and specialized binding-pocket properties, that may have contributed to the loss of ligand-binding function during Orco evolution." We also revised another sentence as follows (lines 522-525): "Overall, we systematically explored the relationships among sequence, structure, and docking-derived function in insect ORs, identified structural features potentially associated with Orco specialization, and provided a framework for testing how OR repertoire evolution may relate to macroenvironmental and lifestyle variation."
It is understandable that the authors want to emphasize the value of their work, but this can't be achieved by inaccurately portraying the state of the field. The framing should be adjusted throughout the manuscript.
We thank the reviewer for this important suggestion. We agree that the manuscript should emphasize its contribution without overstating gaps in the field. We have therefore revised the framing throughout the manuscript to better acknowledge previous advances in insect OR structure, function, evolution, and ecological adaptation. The revised text presents our main contribution as an insect-classscale integration and extension of these findings using a unified sequence, structure, and docking-derived binding-profile framework.
(2) It is unclear how complete the odorant receptor sets are, because the annotation pipeline used is not benchmarked. The high similarity of ORs often located in clusters of up to 50 genes on the genome represents a major obstacle for purely automated annotation, leading to artifacts such as the erroneous joining of exons across genes. The lack of manual verification in this study, combined with stringent filtering of annotations that do not meet structural criteria, does likely lead to a set of sequences that present a correct fold, but at the same time, the approach may discard misannotated genes and cannot distinguish between correct gene models and artifacts where exons of more than one gene could have been merged. Please benchmark the pipeline by comparing its output to a gold standard and frequently vetted complete OR set, such as that of Robertson and Wanner 2006 or similar. That would allow assessing the presented work better and reveal how complete the OR counts are. This is important since the authors use OR counts to derive conclusions about the evolutionary dynamics of the gene family.
We thank the reviewer for raising this concern. Because this point overlaps with the benchmark issue raised in the public review, we provide the detailed benchmark results in our response above and in the revised supplementary text. Briefly, we compared the annotation and structural-filtering pipeline with curated OR repertoires from Drosophila melanogaster, Anopheles gambiae, and Bombyx mori. The results show that the workflow recovers most known ORs, while the structural filter removes fragmented or structurally incomplete models. We also added sensitivity analyses using published OR counts where available, which supported the main OR-count ecological associations.
(3) The in silico functional docking analysis is intriguing and bears the potential to produce testable hypotheses on the functional evolution of ORs. Docking can produce a wide range of results, including artifacts, and thus should be benchmarked with existing datasets derived from functional experiments (there are several large datasets available). While the authors have used previous functional data to constrain their model, they should benchmark it by running it on a set of ORs with known ligand-binding profiles to convincingly show that their predictions are an accurate assessment of the actual functional properties of ORs.
We thank the reviewer for this suggestion. We agree that docking-derived results should be evaluated using ORs with known ligand-response data. In this study, however, the key prediction used for downstream analyses is the dFunC functional label derived from each OR’s overall docking-derived binding profile, rather than the exact ligand assignment of each OR-VOC pair. Therefore, we evaluated the dFunC labels using the Drosophila melanogaster experimental OR response matrix.
Drosophila ORs were mapped to dFunC communities, and for each VOC functional group, ORs were assigned positive or negative labels according to the predicted binding tendency of their dFunC. These labels were compared with experimentally measured response counts from the Drosophila OR functional matrix. ROC curves and AUC values were calculated for each VOC functional group. Most functional-group labels achieved AUC values above 0.75, indicating that the dFunC classification is broadly consistent with the Drosophila experimental response matrix.
We did not use individual OR-VOC pair accuracy as the primary validation criterion because the hit rate analysis showed that the experimental positive rate in the enriched docking-score range is approximately 23%. This indicates that docking is more appropriate for evaluating dFunC-level binding profile tendencies than for exact ligand assignment. We have revised the manuscript to clarify this validation logic. The functional labels of dFunCs should be further tested with experimental response data from more insect species and can be updated as prediction methods improve.
The following details have been added to the revised manuscript (lines 794-798): " Drosophila melanogaster ORs were mapped to dFunCs to evaluate the consistency between dFunC labels and experimental response data. For each VOC functional group, ORs were assigned positive or negative labels according to their dFunC labels, and these labels were compared with response counts from the Drosophila functional matrix to generate ROC curves and AUC values."
(4) The authors conclude that holometabolous and non-holometabolous insects exhibit distinct OR differentiation patterns, stating that sequence diversity "increases progressively with increasing degree of insect order divergence, suggesting a gradual accumulation of sequence variation during insect evolution" (Lines 332-333). If the pattern is indeed gradual, is the phylogenetic splitting of holometabolous vs. non-holometabolous insects not arbitrary? Couldn't it be split at any point along the phylogeny and lead to a similar difference?
We thank the reviewer for pointing out this important logic issue. We agree that, if OR sequence variation accumulates gradually with phylogenetic distance, the difference between holometabolous and non-holometabolous insects should not be interpreted as two completely separate OR differentiation modes.
Our result is better understood as a comparison of extant deep insect lineages. In this comparison, holometabolous insects show higher OR sequence-community diversity than non-holometabolous insects. This pattern is still informative because it suggests that, during long-term evolution, OR repertoires in holometabolous lineages have accumulated broader sequence-community diversity. This may be related to lineage-specific OR duplication and loss, ecological diversification, life-history differences, and differences in repertoire size.
We have revised the manuscript accordingly. We no longer describe the result as two distinct OR differentiation patterns. Instead, we describe it as a difference in OR sequence-community diversity between extant holometabolous and non-holometabolous lineages.
The following revision has been added to the revised manuscript (lines 370-371): "Additionally, extant holometabolous insects showed higher OR sequence-community diversity than non-holometabolous
insects."
(5) Further, if I understand correctly, the assessment of the differentiation patterns seems to rest on total counts of ORs in 'SeqCs', quantified in Figure S9. These counts, however, are not normalized and thus seem influenced by the total number of sequenced genomes belonging to a specific clade in the dataset. These have a heavy bias towards holometabolous species. If this is true, the analyses should be repeated based on normalized counts. OR is this assessment based on the number of 'SeqCs' across the phylogeny? If yes, does this correlate with OR numbers? Please clarify.
We thank the reviewer for pointing this out. We agree that unnormalized SeqC counts can be affected by unequal sampling, because holometabolous insects include more sampled species and more ORs in our dataset.
To address this issue, we added normalized analyses. First, we calculated SeqC diversity at the species level (Fig. S9a). Second, because OR number was significantly correlated with SeqC count (pGLS p-value = 0.001035), we performed equal-OR random sampling (Fig. S9 b). Specifically, we randomly sampled the same number of ORs from holometabolous and non-holometabolous insects, 100 ORs per group, calculated how many SeqCs were covered by the sampled ORs, and repeated this procedure 100 times to obtain the distribution of SeqC diversity under equal OR sampling depth. These analyses showed that holometabolous insects still exhibited higher OR sequence-community diversity after controlling for OR sampling depth. We have revised the manuscript to clarify the calculation and now interpret the result as a normalized difference in sequence-community diversity, rather than as an uncorrected difference in total SeqC number.
We added the following normalized analysis to the revised manuscript (lines 371-379): "Because total SeqC counts may be affected by unequal species sampling and OR numbers, we added normalized analyses. First, we calculated SeqC numbers at the species level and found that holometabolous insects contained more SeqCs on average. Second, because OR number was significantly correlated with SeqC count (P=0.001035), we performed equal-OR random sampling between holometabolous and nonholometabolous insects. Under the same OR sampling depth, holometabolous insects still covered more SeqCs, indicating higher OR sequence-community diversity after controlling for OR number. Therefore, this result supports a genuine difference in SeqC diversity rather than a difference driven by unequal species sampling or OR number. "
(6) Phylogenetic differences in OR evolutionary patterns have been described previously for Paleoptera compared to Neoptera. Are the authors simply picking up on these patterns? Is the holometabolous vs non-holometabolous hypothesis supported when only Neoptera are taken into account?
We thank the reviewer for raising this point. Differences in OR evolutionary patterns between Paleoptera and Neoptera have already been described in previous studies, so it was necessary to test whether our pattern simply reflected this older phylogenetic division.
Following the suggestion, we restricted the comparison to Neoptera and compared Holometabola with non-holometabolous Neoptera. After excluding Paleoptera and earlier-diverging lineages, Holometabola still showed higher OR sequence-community diversity (Fig. S10). Additional normalized analyses indicate that this trend is not explained only by unequal species sampling or OR counts (Fig S9a-b).
Therefore, our result is not simply a rediscovery of the known Paleoptera versus Neoptera difference. Instead, within Neoptera we observe an additional difference in SeqC diversity between Holometabola and non-holometabolous Neoptera.
We added the following comparison of neopteran insects to the revised manuscript (lines 379-382): "The higher SeqC diversity observed in Holometabola was retained when the comparison was restricted to Neoptera, indicating that this pattern is not simply a restatement of previously reported Paleoptera-Neoptera differences."
Minor comments
(1) Line 10: "all the insect orders were found to contain fully functional OR repertoires". It is unclear what the authors mean by this. What is a fully functional OR repertoire? Is it the number of receptors? The number of functional receptors? Given this lack of clarity and the fact that the manuscript does not present functional data on Or binding profiles, this part should be altered.
We have revised this sentence. The intended meaning was that all sampled insect orders include ORs assigned to all six docking-derived functional communities.
(2) Lines 17-18: I am not convinced the presented data explain the adaptive relationship between insect olfactory potential and diverse ecological environments. It suggests that few aspects of insect ecology (larval diet and terrestriality) correlate with receptor number and predicted ligand binding breadth. This should be toned down.
We have toned down the statement. The sentence has been revised as follows: " Our findings provide a class-level framework for investigating insect OR evolution and generate testable hypotheses about how receptor repertoire diversification may relate to ecological adaptation."
(3) Line 44: the low amino acid sequence identity among ORs has been described before. Please add appropriate references.
We have added the requested references.
(4) Lines 52 - 55: The 1:3 stoichiometry is not yet confirmed in vivo - indeed, other work suggests a 2:2 stoichiometry as a possibility. Please adjust this sentence to reflect this uncertainty.
We have revised the sentence to reflect this uncertainty (lines 51-56): Unlike vertebrate G protein-coupled ORs, insect ORs function as ligand-gated ion channels by assembling with the conserved co-receptor Orco, with recent cryo-EM structures supporting a 1:3 OR–Orco heterotetrameric model. However, the exact in vivo stoichiometry remains to be fully resolved, and alternative arrangements such as 2:2 may also be possible.
(5) Lines 94-95: Please cite studies that previously annotated ORs.
We have added the relevant citations.
(6) Lines 99-100: It is well established that hymenopterans have the largest sets of ORs among insects. Please add appropriate references.
We have added the relevant citations.
(7) Lines 101-102: It has been shown before that Odonata have very few ORs. However, there are other insects with even smaller sets of ORs. Please add appropriate references.
We have clarified that the statement refers only to our intact OR dataset. The sentence has been revised as follows: "In our OR dataset, Odonata had the fewest intact ORs, with a mean of 4 (N = 3)."
(8) Lines 109-111: This sentence appears to be illogical. Please revise.
We re-evaluated analyses that depend on OR counts. Because the association between genome size and OR count was sensitive to annotation completeness, structural filtering, and a few extreme species, we removed this result and the related discussion from the revised manuscript.
(9) Line 113: What is ecological behavior? Do you mean ecology?
We have replaced ecological behavior with ecological traits.
(10) Lines 148 - 150: This has been described previously. Please add appropriate references.
We have added the appropriate citation.
(11) Lines 161-162: This has been previously described. Please add appropriate references.
We have added the appropriate citation.
(12) Lines 199-201: Could it be that a lack of orthologs is a result of sampling bias? Only very few genomes of early-branching lineages have been sequenced.
We thank the reviewer for this helpful comment. We agree that the apparent lack of direct orthologs among early-diverging insect ORs may be influenced by limited genome sampling from these lineages. We have revised the manuscript to clarify that, with the currently available data, we cannot reliably determine direct orthologous relationships among early-diverging insect ORs or reconstruct ancestral OR states. We now state that broader sampling of high-quality genomes and transcriptomes from early-diverging insect lineages will be needed to test this question more rigorously.
The revised text is as follows (lines 218-220): "With the currently available early-diverging insect genomes, we could not reliably identify direct orthologous relationships among early ORs, which limits the reconstruction of ancestral OR states."
(13) Line 204-205: Do you mean TdomOR1-8 may be ancestral to Orco? This is not possible, given that Orco and TdomOR1-8 co-exist in one genome. But previous phylogenetic analyses of multiple silverfish genomes suggest that Orco is closely related to TdomOR1-8 and similar receptors in other species. Please revise.
We thank the reviewer for pointing this out. We did not mean that TdomOR1-8 are the ancestors of TdomOrco. Because currently available OR data from early-diverging insects are limited, direct orthologous relationships cannot be reliably identified and the ancestral OR state of Zygentoma cannot be reconstructed with confidence. Therefore, we can only search among extant zygentoman ORs for comparators that are closely related to the Orco clade.
The current gene tree shows that TdomOR1-8 are closely related to TdomOrco. Thus, they can serve as structural references for comparing Orco-related receptors and for understanding structural changes that may have been involved in Orco specialization.
In the revised manuscript, we have changed the relevant sentence to (lines 223-225): “Therefore, we speculate that Orco recruitment in the ancestral zygentoman lineage may have been accompanied by expansion of a set of candidate ORs, and that TdomOR1-8 represent extant retained members of this receptor set.”
(14) Lines 387-388, this was already known. Please cite accordingly
We have added the appropriate citation.
(15) Line 389: Please include a reference for the finding that early ORs have a homologous tetrameric configuration.
We have added the appropriate citation.
(16) Lines 403-407: This hypothesis has been previously put forward by del Marmol and colleagues. Add reference.
We have added the appropriate citation.
Author response table 1.
Number of OR genes in previously annotated species.
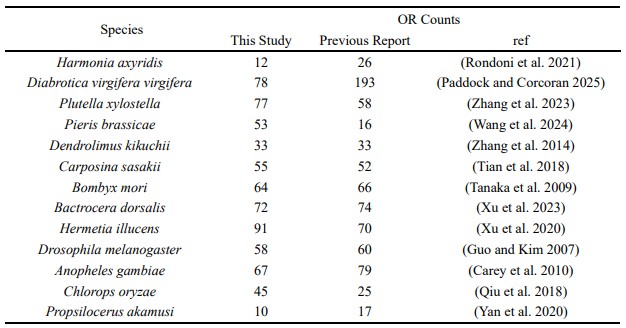
Author response table 2.
ORs in this study that did not show one-to-one correspondence with published Bombyx mori ORs.
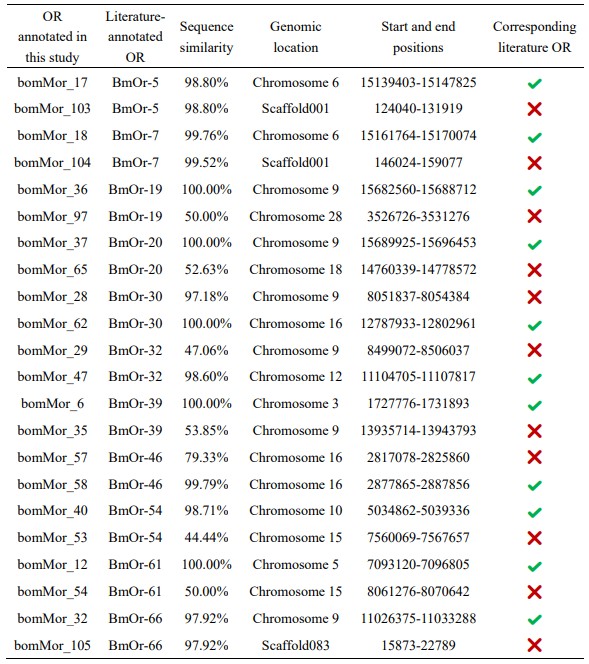
Author response image 1.
Structurally erroneous ORs in Propsilocerus akamusi.
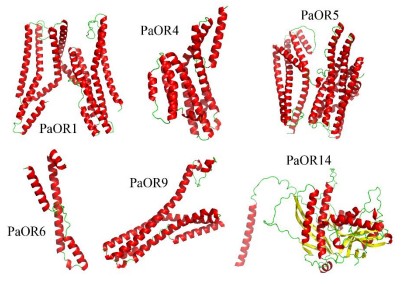
Author response image 2.
Sensitivity analysis of OR counts. (a-b) Correlations between updated species-level OR counts and genome size.
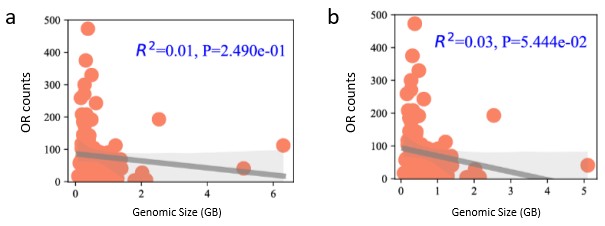
Author response image 3.
OR protein structures removed by structural filtering from the automated annotations of Drosophila melanogaster and Anopheles gambiae.
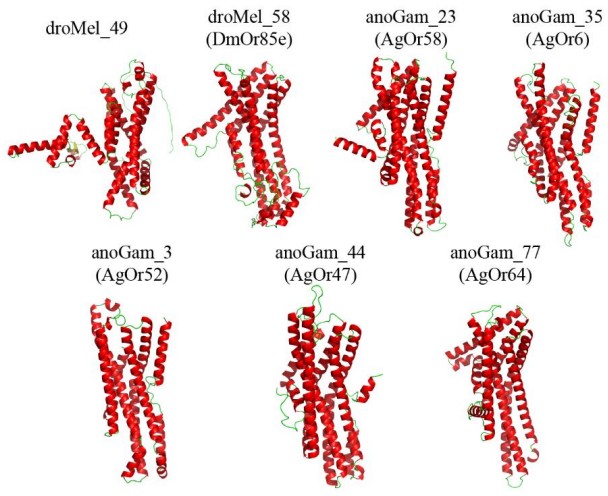
Author response image 4.
Normalized analysis of SeqC diversity between holometabolous and non-holometabolous insects within Neoptera at the species level (a) and OR-count level (b). Welch's t-test was used. ***p < 0.001.
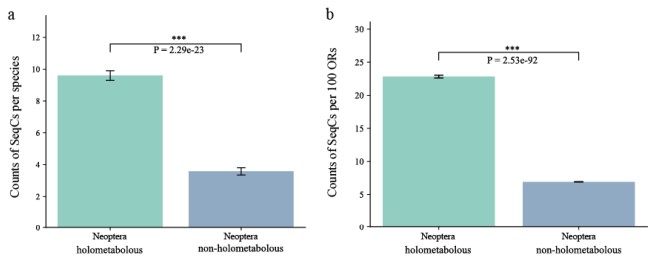
Reference
Bobkov YV, Walker Iii WB, Cattaneo AM. 2021. Altered functional properties of the codling moth Orco mutagenized in the intracellular loop-3. Sci Rep 11: 3893.
Carey AF, Wang G, Su CY, Zwiebel LJ, Carlson JR. 2010. Odorant reception in the malaria mosquito Anopheles gambiae. Nature 464: 66-71.
Chang H, Unni AP, Tom MT, Cao Q, Liu Y, Wang G, Llorca LC, Brase S, Bucks S, Weniger K et al. 2023. Odorant detection in a locust exhibits unusually low redundancy. Curr Biol 33: 5427-5438 e5425.
Dekel A, Pitts RJ, Yakir E, Bohbot JD. 2016. Evolutionarily conserved odorant receptor function questions ecological context of octenol role in mosquitoes. Sci Rep 6: 37330.
Frunze O, Lee D, Lee S, Kwon HW. 2024. A single mutation in the mosquito (Aedes aegypti) olfactory receptor 8 causes loss of function to 1-octen-3-ol. Insect Biochem Mol Biol 167: 104069.
Guo S, Kim J. 2007. Molecular evolution of Drosophila odorant receptor genes. Mol Biol Evol 24: 11981207.
Hall DR, Beevor PS, Cork A, Nesbitt BF, Vale GA. 1984. 1-Octen-3-ol. International Journal of Tropical Insect Science 5: 335-339.
Kline DL, Allan SA, Bernier UR, Welch CH. 2007. Evaluation of the enantiomers of 1-octen-3-ol and 1octyn-3-ol as attractants for mosquitoes associated with a freshwater swamp in Florida, U.S.A. Med Vet Entomol 21: 323-331.
Kreher SA, Mathew D, Kim J, Carlson JR. 2008. Translation of sensory input into behavioral output via an olfactory system. Neuron 59: 110-124.
Liu WB, Li HM, Wang GR, Cao HQ, Wang B. 2023. Conserved Odorant Receptor, EcorOR4, Mediates Attraction of Mated Female Eupeodes corollae to 1-Octen-3-ol. J Agric Food Chem 71: 1837-1844.
Lu T, Qiu YT, Wang G, Kwon JY, Rutzler M, Kwon HW, Pitts RJ, van Loon JJ, Takken W, Carlson JR et al. 2007. Odor coding in the maxillary palp of the malaria vector mosquito Anopheles gambiae. Curr Biol 17: 1533-1544.
Morawo T, Fadamiro H. 2016. Identification of Key Plant-Associated Volatiles Emitted by Heliothis virescens Larvae that Attract the Parasitoid, Microplitis croceipes: Implications for Parasitoid Perception of Odor Blends. J Chem Ecol 42: 1112-1121.
Paddock KJ, Corcoran JA. 2025. Life-stage dependent behavior mimics chemosensory repertoire diversity in a belowground, specialist herbivore. G3 (Bethesda) 15.
Pierce A, Pierce H, Borden J, Oehlschlager C. 1989. Production Dynamics of Cucujolide Pheromones and Identification of I-Octen-3-o1 as a New Aggregation Pheromone for Oryzaephilus surinamensis and O. mercator (Coleoptera: Cucujidae). Environmental Entomology 18: 747-755.
Qiu L, Tao S, He H, Ding W, Li Y. 2018. Transcriptomics reveal the molecular underpinnings of chemosensory proteins in Chlorops oryzae. BMC Genomics 19: 890.
Rondoni G, Roman A, Meslin C, Montagne N, Conti E, Jacquin-Joly E. 2021. Antennal Transcriptome Analysis and Identification of Candidate Chemosensory Genes of the Harlequin Ladybird Beetle, Harmonia axyridis (Pallas) (Coleoptera: Coccinellidae). Insects 12.
Tanaka K, Uda Y, Ono Y, Nakagawa T, Suwa M, Yamaoka R, Touhara K. 2009. Highly selective tuning of a silkworm olfactory receptor to a key mulberry leaf volatile. Curr Biol 19: 881-890.
Tian Z, Sun L, Li Y, Quan L, Zhang H, Yan W, Yue Q, Qiu G. 2018. Antennal transcriptome analysis of the chemosensory gene families in Carposina sasakii (Lepidoptera: Carposinidae). BMC Genomics 19: 544.
Turner RM, Derryberry SL, Kumar BN, Brittain T, Zwiebel LJ, Newcomb RD, Christie DL. 2014. Mutational analysis of cysteine residues of the insect odorant co-receptor (Orco) from Drosophila melanogaster reveals differential effects on agonist- and odorant-tuning receptor-dependent activation. J Biol Chem 289: 31837-31845.
Wang Q, Smid HM, Dicke M, Haverkamp A. 2024. The olfactory system of Pieris brassicae caterpillars: from receptors to glomeruli. Insect Sci 31: 469-488.
Xu L, Jiang HB, Yu JL, Pan D, Tao Y, Lei Q, Chen Y, Liu Z, Wang JJ. 2023. Two odorant receptors regulate 1-octen-3-ol induced oviposition behavior in the oriental fruit fly. Commun Biol 6: 176.
Xu P, Zhu F, Buss GK, Leal WS. 2015. 1-Octen-3-ol - the attractant that repels. F1000Res 4: 156.
Xu Q, Wu Z, Zeng X, An X. 2020. Identification and Expression Profiling of Chemosensory Genes in Hermetia illucens via a Transcriptomic Analysis. Front Physiol 11: 720.
Yan C, Sun X, Cao W, Li R, Zhao C, Sun Z, Liu W, Pan L. 2020. Identification and expression pattern of chemosensory genes in the transcriptome of Propsilocerus akamusi. PeerJ 8: e9584.
Zhang S, Zhang Z, Wang H, Kong X. 2014. Antennal transcriptome analysis and comparison of olfactory genes in two sympatric defoliators, Dendrolimus houi and Dendrolimus kikuchii (Lepidoptera: Lasiocampidae). Insect Biochem Mol Biol 52: 69-81.
Zhang Y, Wang B, Zhou Y, Liao M, Sheng C, Cao H, Gao Q. 2023. Identification and characterization of odorant receptors in Plutella xylostella antenna response to 2,3-dimethyl-6-(1-hydroxy)-pyrazine. Pestic Biochem Physiol 194: 105523.
-
-
-
eLife Assessment
This large-scale comparative study of odorant receptor (OR) genes across more than 100 insect species, combining sequence- and structure-based approaches, aims to explore the evolution of this large gene family involved in the detection of odorant signals by olfactory neurons. This useful work uncovers a structural feature unique to the odorant receptor co-receptor Orco that reduces ligand binding affinity. However, the strength of evidence is incomplete: the pipeline for in silico identification of odorant receptor genes lacks validation through comparison with known odorant receptor repertoires from previously studied species, and claims regarding odor response spectra, evolutionary, and ecological interpretations are not fully supported by the analyses.
-
Reviewer #1 (Public review):
Objectives of the study and impact of the work:
The authors of this article primarily aim to reconstruct the evolutionary history of the insect odorant receptor (OR) family, which is responsible for the detection of odorant signals by olfactory neurons. Due to the lack of phylogenetic signal present in the sequences of this multigene family, which evolves very rapidly, phylogenetic analyses have so far never made it possible to precisely retrace how ORs diversified prior to the appearance of present-day insect orders, and what the drivers of this diversification were. For example, one may suspect that the adaptation of ORs to odors emitted by plants constituted a critical step in insect evolution during the "angiosperm terrestrial revolution," which occurred at the end of the Cretaceous, but nothing …
Reviewer #1 (Public review):
Objectives of the study and impact of the work:
The authors of this article primarily aim to reconstruct the evolutionary history of the insect odorant receptor (OR) family, which is responsible for the detection of odorant signals by olfactory neurons. Due to the lack of phylogenetic signal present in the sequences of this multigene family, which evolves very rapidly, phylogenetic analyses have so far never made it possible to precisely retrace how ORs diversified prior to the appearance of present-day insect orders, and what the drivers of this diversification were. For example, one may suspect that the adaptation of ORs to odors emitted by plants constituted a critical step in insect evolution during the "angiosperm terrestrial revolution," which occurred at the end of the Cretaceous, but nothing currently allows this to be asserted.
There are very nice examples, notably in Drosophilids, derived from comparisons between closely related species and documenting mechanisms of OR adaptation to certain signals. However, what the authors attempt to do in this work is to produce a macroevolutionary analysis at the scale of insects as a whole, based almost exclusively on bioinformatic analyses. To do this, they annotated OR genes in about one hundred insect species and developed pipelines for analyzing sequence similarity, structural similarity, and functional similarity, the latter being estimated through a molecular docking approach. An important feature in the evolution of insect ORs is the emergence of a unique co-receptor, called Orco, which appears to be an OR that has lost the ability to bind odorants. In addition to the large-scale bioinformatic analysis, the authors also aim to explore more specifically the factors that favored the emergence of Orco and the selective advantage conferred by the existence of OR-Orco complexes.
Given the importance of odorant receptors in insect biology and in their adaptation to different environments and lifestyles, retracing their evolutionary history is indeed a major question in evolutionary biology. In principle, this type of work therefore has the potential to become a reference in the field and to provide a basis for significant scientific advances.
Major strengths and weaknesses:
The sampling chosen for collecting OR sequences is very impressive, with more than 100 insect families represented, covering most of the major orders. This sampling appears appropriate for the question being addressed. The analysis pipeline used to collect the sequences makes sense, relying on homology-based annotation tools coupled with a structure-based filter. Nevertheless, one can note aberrant numbers of ORs for certain species (much lower than reality), which indicates that the pipeline probably did not function correctly for all genomes. In the absence of a validation step comparing the results with already known OR repertoires, it is difficult to estimate the overall quality of the data. The authors chose to apply a fairly stringent filter on sequence quality (based on predicted 3D structure), which reduces the number from 14,000 to 9,000. This choice seems logical given the subsequent use of these data, but it inevitably leads to data loss. The fact that some OR genes may be missing and that the total number may not be exact for each species is not prohibitive for studying the evolution of the family at a broad scale; however, it calls into question certain results that rely on this total number, such as the correlation between the number of ORs and genome size, lifestyle, and diet.
From the dataset collected, the authors attempted to categorize ORs in several ways, starting with the reconstruction of sequence similarity networks. The approach is interesting, but in the end, the results do not seem to be sufficiently exploited, and it is not obvious what the advantage of this approach is compared with the "classical" phylogenetic approach, which generally fails to reveal homology relationships between ORs from species belonging to different insect orders. Here again, the majority of the clusters identified are "order-specific," and when this is not the case, the authors did not attempt to exploit the results. For example, clusters SeqC26 or SeqC28, which appear to be shared by many insects, are potentially very interesting. It might have been relevant to combine this similarity-based clustering approach with phylogenetic reconstructions within each shared cluster.
The clustering based on structure also leads to the identification of a majority of "order-specific" clusters, but once again, the clusters shared by several orders are not truly exploited, which does not provide new insight into the evolution of ORs. However, the authors highlight a group of ORs in flies that appear to possess an unusual intracellular region. This is interesting, although it is a result more relevant to OR structure than to their evolution. The function of these ORs in Drosophila melanogaster, if it is known, is not discussed.
The analysis of structural diversity then leads the authors to focus on the Orco co-receptors, which are characterized by modifications of the binding pocket and the emergence of an extracellular loop that could explain the loss of the ability to bind odorant molecules. This part, which relies on in vitro experiments, is interesting and constitutes the most striking result of the study, which could in itself have been the subject of a separate manuscript. However, the molecular dynamics modelling does not add anything in the way it is conducted (5 ns is too short).
The rest of the manuscript is based on the prediction of OR response spectra using molecular docking. The work that has been carried out is extremely substantial, and the objective of linking clusters based on sequence similarity or 3D structural similarity with functional categories is entirely relevant. Nevertheless, I see two major problems with this in silico functional analysis:
(1) The docking score threshold used was chosen thoughtfully, which is very good, and according to the calculation performed, should ensure a true positive rate of more than 20%, which is excellent in such a docking analysis. But in the absence of functional validation, this 20% true positive rate is not sufficient to extrapolate OR function as the authors do in the remainder of the manuscript. The risk of error remains too high to compare in such detail the function of ORs from insects with different lifestyles or diets.
(2) The six functional clusters identified are only slightly different from one another, with similar detection of all chemical families except acids and amines (which was expected, given that these families are a priori detected by IRs rather than ORs). This shows that even though the approach is relevant and deserves to be tested, it cannot be used to establish a link between groups/lineages of ORs and response spectra at the scale of insects as a whole. This is reflected in the final analysis by the fact that there is no visible link between sequence or structural clusters and functional clusters. Given the uncertainty surrounding the docking results, the entire subsequent analysis of the relationship between the Binding Breadth Index and ecological variables is highly questionable.
Finally, the evolutionary analysis proposed to conclude that the work suffers from an incorrect interpretation: ORs of non-holometabolous insects cannot be considered equivalent to those of species that existed before the Permian-Triassic extinction. The fact that a locust or a cockroach has more narrowly tuned ORs than holometabolous insects does not mean that this was also the case for ancestral insects. To advance this type of conclusion, it would be necessary to conduct a phylogenetic analysis and reconstruct ancestral states, which is not the case here.
In summary, despite the large number of analyses performed, the authors do not succeed in achieving the stated objective of reconstructing the evolutionary history of insect ORs, and the results obtained do not sufficiently support the conclusions regarding the links between OR repertoires and environment or lifestyle.
-
Reviewer #2 (Public review):
The remarkable evolvability of the olfactory system enables animals to rapidly adapt to dynamic and chemically complex environments. Over the past two decades, substantial effort has been devoted to uncovering the evolutionary principles that drive the diversification of odorant receptors (ORs), yielding key insights into the forces shaping their striking variability in both vertebrates and insects. In this manuscript, Zhang and colleagues analyze the OR repertoires of over 100 insect species, leveraging sequence and structural similarity to infer patterns of gene family evolution within this diverse and ecologically important clade. By integrating sequence-based and structure-based comparisons, their study builds on a compelling and recently emerging line of research made possible by the advent of …
Reviewer #2 (Public review):
The remarkable evolvability of the olfactory system enables animals to rapidly adapt to dynamic and chemically complex environments. Over the past two decades, substantial effort has been devoted to uncovering the evolutionary principles that drive the diversification of odorant receptors (ORs), yielding key insights into the forces shaping their striking variability in both vertebrates and insects. In this manuscript, Zhang and colleagues analyze the OR repertoires of over 100 insect species, leveraging sequence and structural similarity to infer patterns of gene family evolution within this diverse and ecologically important clade. By integrating sequence-based and structure-based comparisons, their study builds on a compelling and recently emerging line of research made possible by the advent of AlphaFold, which has previously clarified the phylogenetic relationship between insect Ors and the gustatory receptor gene family and revealed the unexpectedly deep evolutionary origins of this ancient structural fold.
Applying this approach to a large set of ORs derived from species throughout the insect phylogeny, the authors confirm many previously reported patterns of OR evolution. Unfortunately, the way these results are presented lacks clarity in what is already known from previous work in the field versus what is a novel finding based on the analysis of this dataset.
It is unclear how complete the odorant receptor sets are. I recommend benchmarking the pipeline by comparing its output to a gold standard and a frequently vetted complete OR set, such as that of Robertson and Wanner 2006 or similar.
Using their structural clustering approach, the authors identify a structural feature mostly unique to the OR co-receptor ORco, a beta-sheet in EL2, which they functionally show reduces odorant binding affinity - a key aspect of ORco, which does not bind ligands in the ancestral ligand-binding site. This is a particularly strong part of the manuscript, since the authors support their in silico-derived hypothesis with functional data.
Lastly, in an attempt to assess the relationship between sequence identity and structure on one hand and function on the other, the authors perform an in silico structure prediction and chemical docking analysis. As it stands, this part is on the more speculative side since the docking approach has not been verified with available functional datasets.
-
The authors included RRIDs in their in bioRxiv paper! Thanks for making your methods matter! #OpenScience #methodsmatter #STMpublishing
-
Evolutionary dynamics of insect odorant receptors reveal ecological tuning shaping olfactory perception https://www.biorxiv.org/content/10.64898/2026.02.12.705626v1
-
Evolutionary dynamics of insect odorant receptors reveal ecological tuning shaping olfactory perception https://www.biorxiv.org/content/10.64898/2026.02.12.705626v1
-
Evolutionary dynamics of insect odorant receptors reveal ecological tuning shaping olfactory perception https://www.biorxiv.org/content/10.64898/2026.02.12.705626v1
-